Working to Create Rare Disease Advisory Councils in all 50 States

Fifteen states done, 35 to go.

Fifteen states have established a Rare Disease Advisory Council (RDAC) to give the rare disease community, including those living with Leber congenital amaurosis (LCA), a stronger voice in state government. Another dozen states are actively working to establish RDACs this year.

With assistance from the National Organization for Rare Disorders (NORD), patient organizations, such as Hope in Focus (formally Sofia Sees Hope), and the broader rare disease community, RDACs around the country work to help states strategically address barriers faced by people living with rare diseases. The councils give stakeholders an opportunity to make formal recommendations to state leaders on critical rare disease issues, including increased awareness, diagnostic tools, and access to affordable treatments and cures.

Sofia Sees Hope, based in Ledyard, Conn., annually gives information to legislators on the Connecticut General Assembly’s Public Health Committee, letting them know that rare disease advocates and those living with a rare disease, need state and federal support in crafting legislation to help the rare disease community. 

The last day of February each year is officially Rare Disease Day, a time to raise awareness among the public and decision makers about rare diseases and their impacts on patients’ lives. NORD sponsors Rare Disease Day in the United States, alongside its sister organization, the European Organization for Rare Disorders (EURODIS), which organizes the official international campaign.

Find out here what is happening globally in celebration of Rare Disease Day. 

7,000 Rare Diseases in the U.S.

A disease is defined as rare in the United States if it affects fewer than 200,000 Americans. LCA and other inherited retinal diseases (IRDs) are among the approximately 7,000 rare diseases that exist nationally, affecting 1 in 10 people. These diseases include more than 500 types of rare cancers and all pediatric cancers. 

Between 25-30 million Americans live with a rare disease, including about 300,000 in Connecticut. 

NORD’s Connecticut Rare Action Network, along with other such groups nationwide, will highlight RDACs as part of its virtual celebration of Rare Disease Day 2021. Connecticut’s Rare Disease Day celebration will be at 9 a.m. ET, Friday, Feb. 26. You can register to participate in this free event by clicking here. 

NORD also just released its sixth edition of its State Report Card* that analyzes the 50 states and Washington, D.C., on eight policy issues that impact the rare disease community. The organization launched its report card project to evaluate the effectiveness of states serving people with rare diseases.

Each state has its own report card that addresses Medicaid Financial Eligibility, Medicaid Nutrition, Newborn Screening, Prescription Drug Out-of-Pocket Costs, Protecting Patients in State Medicaid Programs, Protecting Patients in State-Regulated Insurance, Rare Disease Advisory Councils, and Step Therapy (trying less expensive options before “stepping up” to drugs that cost more).

State governments make decisions every day affecting the rare community. They play critical roles in ensuring access to health care providers, services, and treatments needed to thrive, along with the design of their Medicaid program benefits, and regulation of some insurance plans. The councils offer forums for discussion about these issues.

Learn which states have Rare Disease Advisory Councils

As one Connecticut legislator said at a previous celebration, having one day to recognize rare diseases is not enough. 

“It needs to be Rare Disease Day every day in the state of Connecticut,” said State Rep. Jonathan Steinberg, co-chairman of the General Assembly’s Public Health Committee.

Connecticut Volunteer State Ambassador Lesley Bennet  – along with ambassadors nationwide, doctors, researchers, advocates, caregivers, patients, legislators, and business leaders – take this time to advocate state-by-state for better resources and outcomes for people living with rare diseases.

Bennett said many of their patients have difficulty getting access to services because people don’t understand the disorders. 

Patients, caregivers, families, and providers in North Carolina created the first RDAC six years ago; 15 states have active councils with1 2 more on their way toward fruition, for a total of 27 states.

Another 23 states, including Connecticut, do not have such councils but efforts continue through NORD’s Project RDAC, launched last fall. Connecticut established a task force to study whether to create an RDAC, but the group disbanded in 2019. 

Check out this map to see if your state has an RDAC or if NORD is working on legislation to create one. To start an effort in a new state, go to RDAC@rarediseases.org. 

Council composition varies from state to state in size, duties, and accountability requirements. It also depends on the type of entity that houses the RDAC, such as a state department of health or a non-profit organization.

A council typically comprises various stakeholders, including patients, caregivers, doctors, insurers, biotechnology companies, researchers, and state officials.

Project RDAC aims to optimize the existing councils and increase the number of groups nationwide by opening collaborative opportunities among the councils, creating educational resources to guide them, and helping more states pass legislation establishing high-functioning councils.
Which States Have Rare Disease Advisory Councils?

15 states with an established Rare Disease Advisory Council:

Alabama, Kentucky, Illinois, Massachusetts, Minnesota, Missouri, Nevada, New Hampshire, New York, North Carolina, Ohio, Pennsylvania, Utah, Tennessee, and West Virginia.

Arkansas, California, Florida, Georgia, Kansas, Michigan, New Jersey, South Carolina, Texas, Virginia, Washington, and Wisconsin.

23 states with no Rare Disease Advisory Council:

Alaska, Arizona, Colorado, Connecticut, Delaware, Hawaii, Idaho, Indiana, Iowa, Louisiana, Maryland, Maine, Mississippi, Montana, Nebraska, New Mexico, North Dakota, Oklahoma, Oregon, Rhode Island, South Dakota, Vermont, and Wyoming.

‘Let’s Chat About …’ Webinar Offers LCA Overview and Updates on Clinical Trials

In the debut of Hope in Focus (formally Sofia Sees Hope) ‘Let’s Chat About …’ monthly webinar series, Ben Shaberman of the Foundation Fighting Blindness, provided his Zoom audience with a plethora of information about Leber congenital amaurosis (LCA), highlighting some of the more than 40 clinical trials underway to find treatments and cures for LCA and other rare inherited retinal diseases (IRDs) and giving updates on promising preclinical research. 

The recorded webinar aired 1 p.m. Wednesday, Jan. 27, 2021, and can be seen here. Elissa Bass, our marketing and communications director, moderated the session.

Shaberman, Senior Director, Scientific Outreach & Community Engagement, stumbled across a science writing position at the Foundation Fighting Blindness 16 years ago without a clue about retinas or blindness. He called his move to the Foundation serendipitous. He knew he made the right choice after hearing retinal researcher Dean Bok, PhD, tell attendees at a 2005 Foundation conference how he was drawn to the field by the seduction of the retina’s myriad complexities and inner workings.

Shaberman, too, felt pulled by the intriguing science of the retina.

As such, so are the 27 forms of LCA that cause varying kinds of visual impairment within each gene mutation and within each affected person. An estimated 8,000 people in the United States have LCA.

The path of retinal research

Shaberman took his audience from the beginnings of identifying the RPE65 gene in 1993 and learning shortly thereafter it could lead to LCA, to using mice models and later studying Briard dogs that had the same gene mutation that caused LCA in humans. A clinical trial at Children’s Hospital of Philadelphia led to the 2017 FDA approval of the breakthrough gene therapy LUXTURNA®, developed by Spark Therapeutics. The drug successfully improved the vision of many of the LCA2-RPE65 patients who received the treatment through subretinal injections.

When children receive an LCA diagnosis, their families should find a good retinal specialist, get regular exams, and ultimately get a confirmed genetic diagnosis to be on the path to more specific information and research into that form of LCA, Shaberman said.

Families also should register with the Foundation’s My Retina Tracker®, a free and secure online registry that facilitates getting a confirmed genetic diagnosis by making registrants eligible for free genetic testing.

The registry becomes your personal retinal health record, updated by you. It employs state-of-the-art database technology to protect privacy and adheres to the highest standards of confidentiality and ethics. 

It also notifies registrants of clinical trials and gives researchers access to their disease data – not their personal information – to advance research and therapy development associated with LCA and IRDs. 

Reading research publications and attending events sponsored by the Foundation and by Sofia Sees Hope also provide opportunities for families to interact and learn the latest research. Shaberman and Bass encouraged people affected by LCA and their families to contact them, respectively, through the Foundation’s website and/or the Sofia Sees Hope website for specific information on clinical trials or other questions and concerns about living with LCA. 

“Yes, it’s work,” Shaberman said. “You have to be your own advocate and your own child’s advocate, but more and more information is becoming available, and that’s the good news.”

Shaberman also reviewed some of the more than 40 retinal clinical trials in the pipeline for LCA and other IRDs:

Join us Feb. 16

February’s “Lets Chat About …” webinar airs at 3 p.m. ET, Tuesday, Feb. 16. Our guest will be Wiley A. Chambers, MD, Supervisory Medical Officer for the Office of New Drugs, Center for Drug Evaluation and Research at the U.S. Food and Drug Administration. Register here.

Reflecting on the Trajectory of IRD Research

When I joined the Foundation Fighting Blindness as a science writer in 2004, I really didn’t know what I was getting into. I knew nothing about the retina, let alone the complex and diverse world of rare inherited retinal diseases (IRDs) that includes Leber congenital amaurosis (LCA). But the research for treatments was cutting-edge and compelling, so I was excited to dive in and learn.

My early assignments were writing about laboratory studies coming out of academic labs. There were virtually no companies in the IRD space and only one or two clinical trials underway for emerging therapies. But there were a lot of studies of genetically engineered mice and rat models of IRDs for gaining a better understanding of disease pathways and testing potential treatments.

Truth be told, I often wondered if and when rodent-tested therapies were really going to make it into human studies and out to the people losing vision. But the scientists conducting the research were mind-blowingly smart and innovative, so I figured they knew what the heck they were doing. With a master’s degree in poetry, who was I to judge?

Fast forward about four years: I was in my hotel room in Fort Lauderdale – there for the annual Association for Research in Vision and Ophthalmology conference – when my manager called and told me three research groups just reported vision improvements in young adults treated with RPE65 gene therapies in Phase 1/2 clinical trials. That was the breakthrough we’d all been waiting for.

People, rather than animals, with severe vision loss were now seeing significantly better. It was the first time an IRD treatment had worked in humans. I will never forget the headline for the article I immediately wrote: “Now They See.” (Note: One of those RPE65 gene therapies later became LUXTURNA®, the first FDA-approved treatment for the eye or an inherited condition.)

After many years of painstaking work, our hope for treatments and cures had finally begun evolving into promise.

There have been several other aha! moments in the ensuing years, but I distinctly recall cathartic encounters at the 2019 American Society of Retinal Specialists in Chicago. As I perused the snack table during breaks (the accomplished snacking professional that I am), several representatives from biotechs developing IRD therapies – companies I’d never even heard of – came up to introduce themselves to me and tell me about their emerging IRD treatments. They didn’t know me or my role, nor had I previously known them; they were just eager to connect with someone from the Foundation Fighting Blindness to get on our radar screen.

I realized then I couldn’t keep track of all the companies (dozens) focused on IRDs and clinical trials (40-plus) underway for potential IRD treatments. But being overwhelmed felt incredibly good, and it meant more good news likely was on the horizon for saving and restoring vision.

While mouse studies are as critical as ever, I can’t remember the last time I wrote an article about one. That’s because most of my writing is now dedicated to reporting on advances, including encouraging vision improvements, being made in human studies.

Make no mistake: Much more work needs to be done before we eradicate the myriad IRDs affecting millions of people across the globe. And, of course, we cannot get more therapies across the finish line fast enough. But when I look at how incredibly far we’ve come since those early days of mice and rats, I have no doubt we are well on our way to breaking many more ribbons soon.

Nearly Three Years Later, LUXTURNA®™ Treatments Continue to See Success

Since its launch in March 2018, breakthrough gene therapy LUXTURNA®™ continues to be successful in helping improve vision in people with inherited retinal disease due to mutations in both copies of the RPE65 gene and viable retinal cells as determined by a healthcare professional. The therapy treats LCA2, known as LCA/RPE65, one of more than 25 forms of Leber congenital amaurosis. 

The drug – developed by Spark Therapeutics and a team of retinal research superstars that included Dr. Katherine A. High  and Dr. Jean Bennett – came to fruition after 12 years of research and millions of dollars in investment. 

Spark Therapeutics could not comment on the number of people who have received the gene therapy, but spokesman Kevin Giordano recently said the company has shipped well over 200 vials of the therapy since the U.S. Food & Drug Administration approval in December 2017. One vial of the drug treats one eye.

Trained retinal surgeons at one of the 10 eligible treatment centers in the United States deliver the gene therapy to the back of the eye by subretinal injection using a needle the size of an eyelash; about a week or so later, the patient undergoes the procedure in the other eye.

Read: 10 US Centers Offering Treatment

“Spark Therapeutics is thrilled patients continue to benefit from LUXTURNA (voretigene neparvovec-ryzl),” Giordano, Spark’s External and Product Communications Lead, said. “A gene therapy is a major milestone, not only for those of us at Spark, but also for the patient community and broader gene therapy field.”

The cost of the drug – $425,000 for each eye – initially caused anxiety among patient families, but Giordano had good news about insurance coverage.

“Payer coverage has exceeded expectations, and to our knowledge no treatment-eligible patient has been denied treatment due to their insurance coverage,” he said.

LUXTURNA also is being used beyond this country through license and supply agreements with Novartis, which has the rights to commercialize the drug in Europe and all other markets outside the United States. Spark manufactures and supplies the gene therapy to Novartis, according to Giordano.

Also, results from ongoing studies continue to support the drug’s safety profile and efficacy.

“In 2019, Spark presented four-year durability data from the LUXTURNA Phase 3 clinical trial,” Giordano said. 

Read: LUXTURNA safety information

For privacy reasons, Spark Therapeutics cannot share patient results.

But we can – at least a couple of them because they came from patients and families sharing their stories with us.

For Hannah, ‘Hope Realized’

Hannah Reif, daughter of Amy and Chris Reif of Maple Glen, PA, underwent LUXTURNA gene therapy in July 2018. Dr. Albert M. Maguire, who is married to researcher Dr. Bennett, performed Hannah’s surgery at Children’s Hospital of Philadelphia (CHOP).  

Hannah with a big smile, wearing a light blue green coat and holding a drink
Hannah Reif

“We closely watched the clinical trials and the FDA approval process for seven years, starting when Hannah was diagnosed with LCA/RPE65 at just a few months of age,” Amy said. “Seven years of hope. 

“Two years out from Hannah’s surgery, I can say we feel grateful and fortunate that she was treated with LUXTURNA. No regrets. LUXTURNA was hope realized. It delivered what it promised.”

She said what that has meant for Hannah has been nothing short of life changing.

“It has given her more independence, which has been wonderful for her self-confidence. It has given her the ability to see what she couldn’t before.”

Since the surgery, Hannah’s vision in dim lighting and her visual acuity improved. She is now 9 and just finished third grade. 

Sometimes, her mom said, it’s the little things that are the most amazing.

“A year after the surgery, she was about to eat hot oatmeal and said, ‘Hey, I see steam. Hey, I can see that,’ ” Amy said.

“There are still things that pop up that she’s seen for the first time, like when she said, ‘Mom, did you know there’s a pattern on this toy?’ It’s fun to see her discovering.”

Amy said she and her family will be forever grateful to Dr. Bennett.

“There has been a lot of talk about heroes recently and Dr. Bennett is our hero. We are grateful for this groundbreaking treatment that she developed, that has been life-changing, not only for our daughter, but also for the sons and daughters and loved ones of so many others as well.”

‘His Vision Changed Everything’

Creed Pettit, one of the first recipients LUXTURNA, received the gene therapy at age 9 in March 2018. Dr. Audina M. Berracol  performed the surgery at Bascom Palmer Eye Institute in Miami, Fla.

His mom, Sarah St. Pierre Schroeder, told us that her now-11-year-old is doing amazing and only occasionally has issues with dim lights, “but nothing like before.”

Their days in Mount Dora, Fla., have changed in a major way.

“He still starts his day with his trusty smoothie and waffle, but after that, Creed wants to create new pranks (today was putting ice in the tub). He said it was nice and warm so I could soak my feet.”

Creed in blue glasses, holding his Uno Cards
Creed Pettit

Creed now loves to play board games – Trouble, Sorry, Battleship and Uno. 

“His vision has changed everything. He can manipulate small objects, he is using pointer fingers, and loves trying to roll his eyes in the mirror.”

Creed still likes to draw, and he loves riding and popping wheelies on his bike named Carlitos. He also likes to talk.

“Talking more is an understatement! Sunup to sundown, he is talking. He has also learned how fun it is to use his imagination, something he never did before. He creates awesome stories during this time.  He is so much more independent; I have to remind myself of that often.”

At first, she and her husband, Chad, could see that Creed’s vision improved some, but once he became confident about his gene replacement, they noticed him finding toys and games.

“He was suddenly enjoying things he didn’t before. He now expresses when he can’t see. Yesterday it rained all day, I kept waiting for him to tell me it was too dark inside, but he was fine. He just started doing staring contests. I love looking at his eyes during the contest.”

Sarah said she is incredibly grateful.

“Creed’s surgery is something we still thank God, St. Raphael, St. Lucy and Sister Miriam Teresa, for every night.” 

Like many kids across the country, Creed finished school at home because of the coronavirus pandemic. He graduated from fifth grade and thought because he finished school at home, he wouldn’t have to wear the graduation cap.

“But I was not going to let that slide,” his mom said. “I made one and took pictures.”

Throughout his journey, though, Creed felt anxious, something Sarah had learned that might happen when, before Creed’s surgery, she talked with Tami Morehouse, a LUXTURNA clinical trial participant at age 44.

Tami, a Sofia Sees Hope Ambassador, tries to calm fears and advises potential gene therapy patients and their parents that even though undergoing the surgery has the potential to do such good by improving vision, they should think about their expectations, especially with children.

“We are comfortable in our own zone; give us a little bit of change and it can throw us off,” Tami said.

Sarah understands. 

“He started to have a lot of anxiety. He had a hard time sleeping. I feel everything changed so fast that he was overwhelmed, but we have worked hard on getting past that.”

Sarah and her family are in a great place now. But after they were home from the hospital in Miami and settled into their routine, she said she became very emotional.

“All the tears I had held in for nine years started to come out. I felt I no longer had a purpose; I was so used to staying busy. I did not know who I was supposed to be.” 

She got some help and realized she still was needed because people need her help learning about and understanding this groundbreaking gene therapy.

“I still find myself shocked over how this has changed Creed’s life and so many other lives.”

Clinical Trials and Emerging Research Show Promise for LCA Treatments

Forty clinical trials and a lot of pre-clinical research into LCA treatments show promising pathways to discovering the next LUXTURNA®, according to Shannon Boye, PhD, the opening speaker for the Virtual VISIONS 2020 conference, presented earlier this summer by the Foundation Fighting Blindness. 

The breakthrough drug developed by Spark Therapeutics marked a milestone in the history of genetic research as the first gene therapy in the United States for any inherited disease and as the first to treat one of the more than 25 forms of Leber congenital amaurosis (LCA).

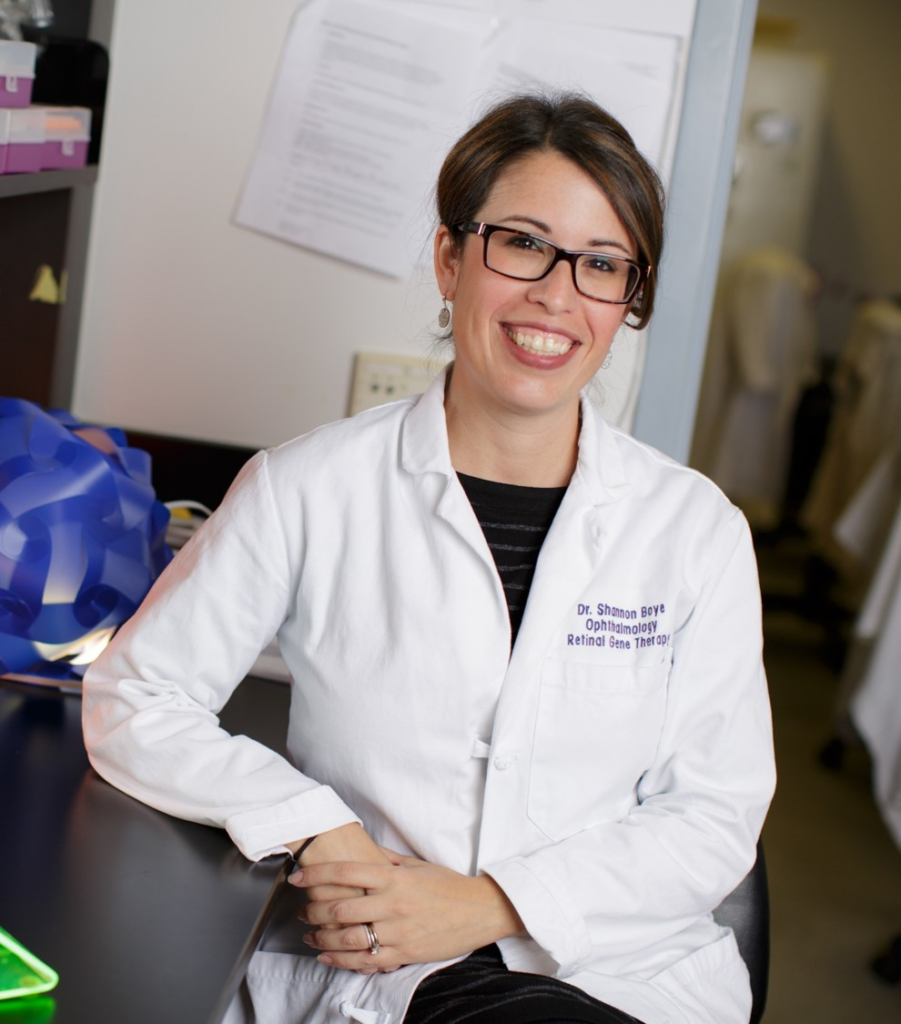
Shannon Boye, PhD in a lab coat
Shannon Boye, PhD

Boye, along with Foundation Chairman of the Board David Brint and Foundation Chief Executive Officer Benjamin Yerxa  PhD, kicked off the three-day, first-time virtual conference, the Foundation’s major annual gathering. Rather than convening in person, the event’s speakers, exhibitors and more than 1,600 attendees participated through an online app, due to concerns surrounding the coronavirus pandemic.

Brint said that the 40 clinical trials and more emerging treatments for various IRDs span the disease profile.

“No matter what your disease is, these hopefully will be able to restore vision,” Brint said. “In the next 10 years, we have an opportunity to bring many more vision-saving treatments into and through the pipeline and across the finish line.”

Yerxa said the topic of genetic therapies would be good to lead off with because of the sheer variety of innovative programs and approaches to each therapeutic challenge.

“There’s essentially a revolution happening right now in personalized medicine and genetic therapies in general,” Yerxa said.

Boye, an assistant professor in the Department of Ophthalmology at the University of Florida, addressed the audience in the beginning session called: “Mission Possible! What’s Next?”

She discussed three major strategies in treating LCA and other IRDs: Gene supplementation or gene replacement therapy, RNA therapeutics and gene editing. 

Boye set up an analogy to better understand the complexities of these strategies, saying we all have little letters in our bodies called DNA. Subunits of those letters – that DNA – are genes. RNA carry the instructions from DNA for making proteins, the building blocks of life.

“They act alone or in concert with a bunch of other proteins to perform essential functions.”

Continuing her letters analogy, Boye said, imagine a friend texts you: ‘Please take out the dog.’ You get that message and perform that function because letters combined correctly to tell you to take the dog out. 

If only the word ‘Please’ appears on your phone screen, you don’t take the dog out.

Or, if the ‘d’ is pushed and an ‘l’ comes out, sending the message, ‘Please take the log out,’ “you then have a mess to clean up,” she quipped.  

In the first strategy of gene supplementation or gene replacement, the right protein needs to be expressed in the patient’s retina. 

The letters need to be correctly sequenced to generate a coherent message, in this case, telling a protein to perform an important function. Any break in that cascade of events can cause visual impairment. 

The gene replacement therapy LUXTURNA is a human-engineered virus containing copies of the corrective gene that doctors deliver through a subretinal injection so the cells can make the originally missing protein.

“You deliver the right letters that make the right message and the right protein,” she said. “That’s a pretty simple concept. That’s LUXTURNA.”

Developed to improve vision in people with LCA2* caused by a mutation in the RPE65 gene, LUXTURNA received Food and Drug Administration approval for  use in humans in December 2017. 

One area of Boye’s research as Associate Division Chief of Cellular and Molecular Therapeutics is entering into a Phase 1/2 clinical trial, applying the same premise for mutations in the GUCY2D gene that causes LCA1. 

“It’s early,” she said. “But this is an example of another perhaps next LUXTURNA being right around the corner.”

She cited similar research moving forward on other IRDs, including Retinitis Pigmentosa (RP), Choroideremia, and Bardet Biedl Syndrome (BBS). 

The second strategy is a form of RNA therapeutics that uses antisense oligonucleotides (AONs) – short, single-stranded DNA  molecules that interact with messenger RNA to correct translation of a targeted gene. Think of an AON as an autocorrect feature that binds to the ‘l’ in log and changes to a ‘d’ for dog.

Promising pre-clinical work now in Phase 2/3 for CEP290 or LCA10 also is coming out of Rob Collin’s research group in The Netherlands, Boye said.

Another AON program underway addresses a form of Usher Syndrome. 

The third strategy – the newest and most exciting – is gene editing. A guide RNA is used to drag a special enzyme to a region in the DNA that contains the mutation, and the enzyme cuts the DNA, like molecular scissors. 

Researchers are exploring a host of gene editing variations, including cutting out a specific area of DNA and replacing it with the right letters to make a coherent message. The lab work has created paths to address a range of IRDs, including CEP290,  Usher Syndrome,  RP, Stargardt Disease and Choroidermia.

“There’s an absolute exponential increase in the therapies that are being developed,” she said.

These strategies are not limited to the disease conditions under discussion and can be more widely applied to a number of genes and conditions.

Addressing those who do not have RPE65 or LCA2 for which a treatment exists, Boye said, with all of this research in progress, “that one day, there’s going to be a LUXTURNA for your inherited retinal disease, too.” 

Rare Disease Day 2020: Lawmakers Hear From Advocates

Rare disease patients, caregivers, advocates, researchers, doctors, healthcare providers and lawmakers gathered at Connecticut’s capitol in Hartford on Friday, Feb. 28, to celebrate Rare Disease Day 2020 and raise awareness because rare medical conditions often are overlooked by health-policy decision makers and the medical community. 

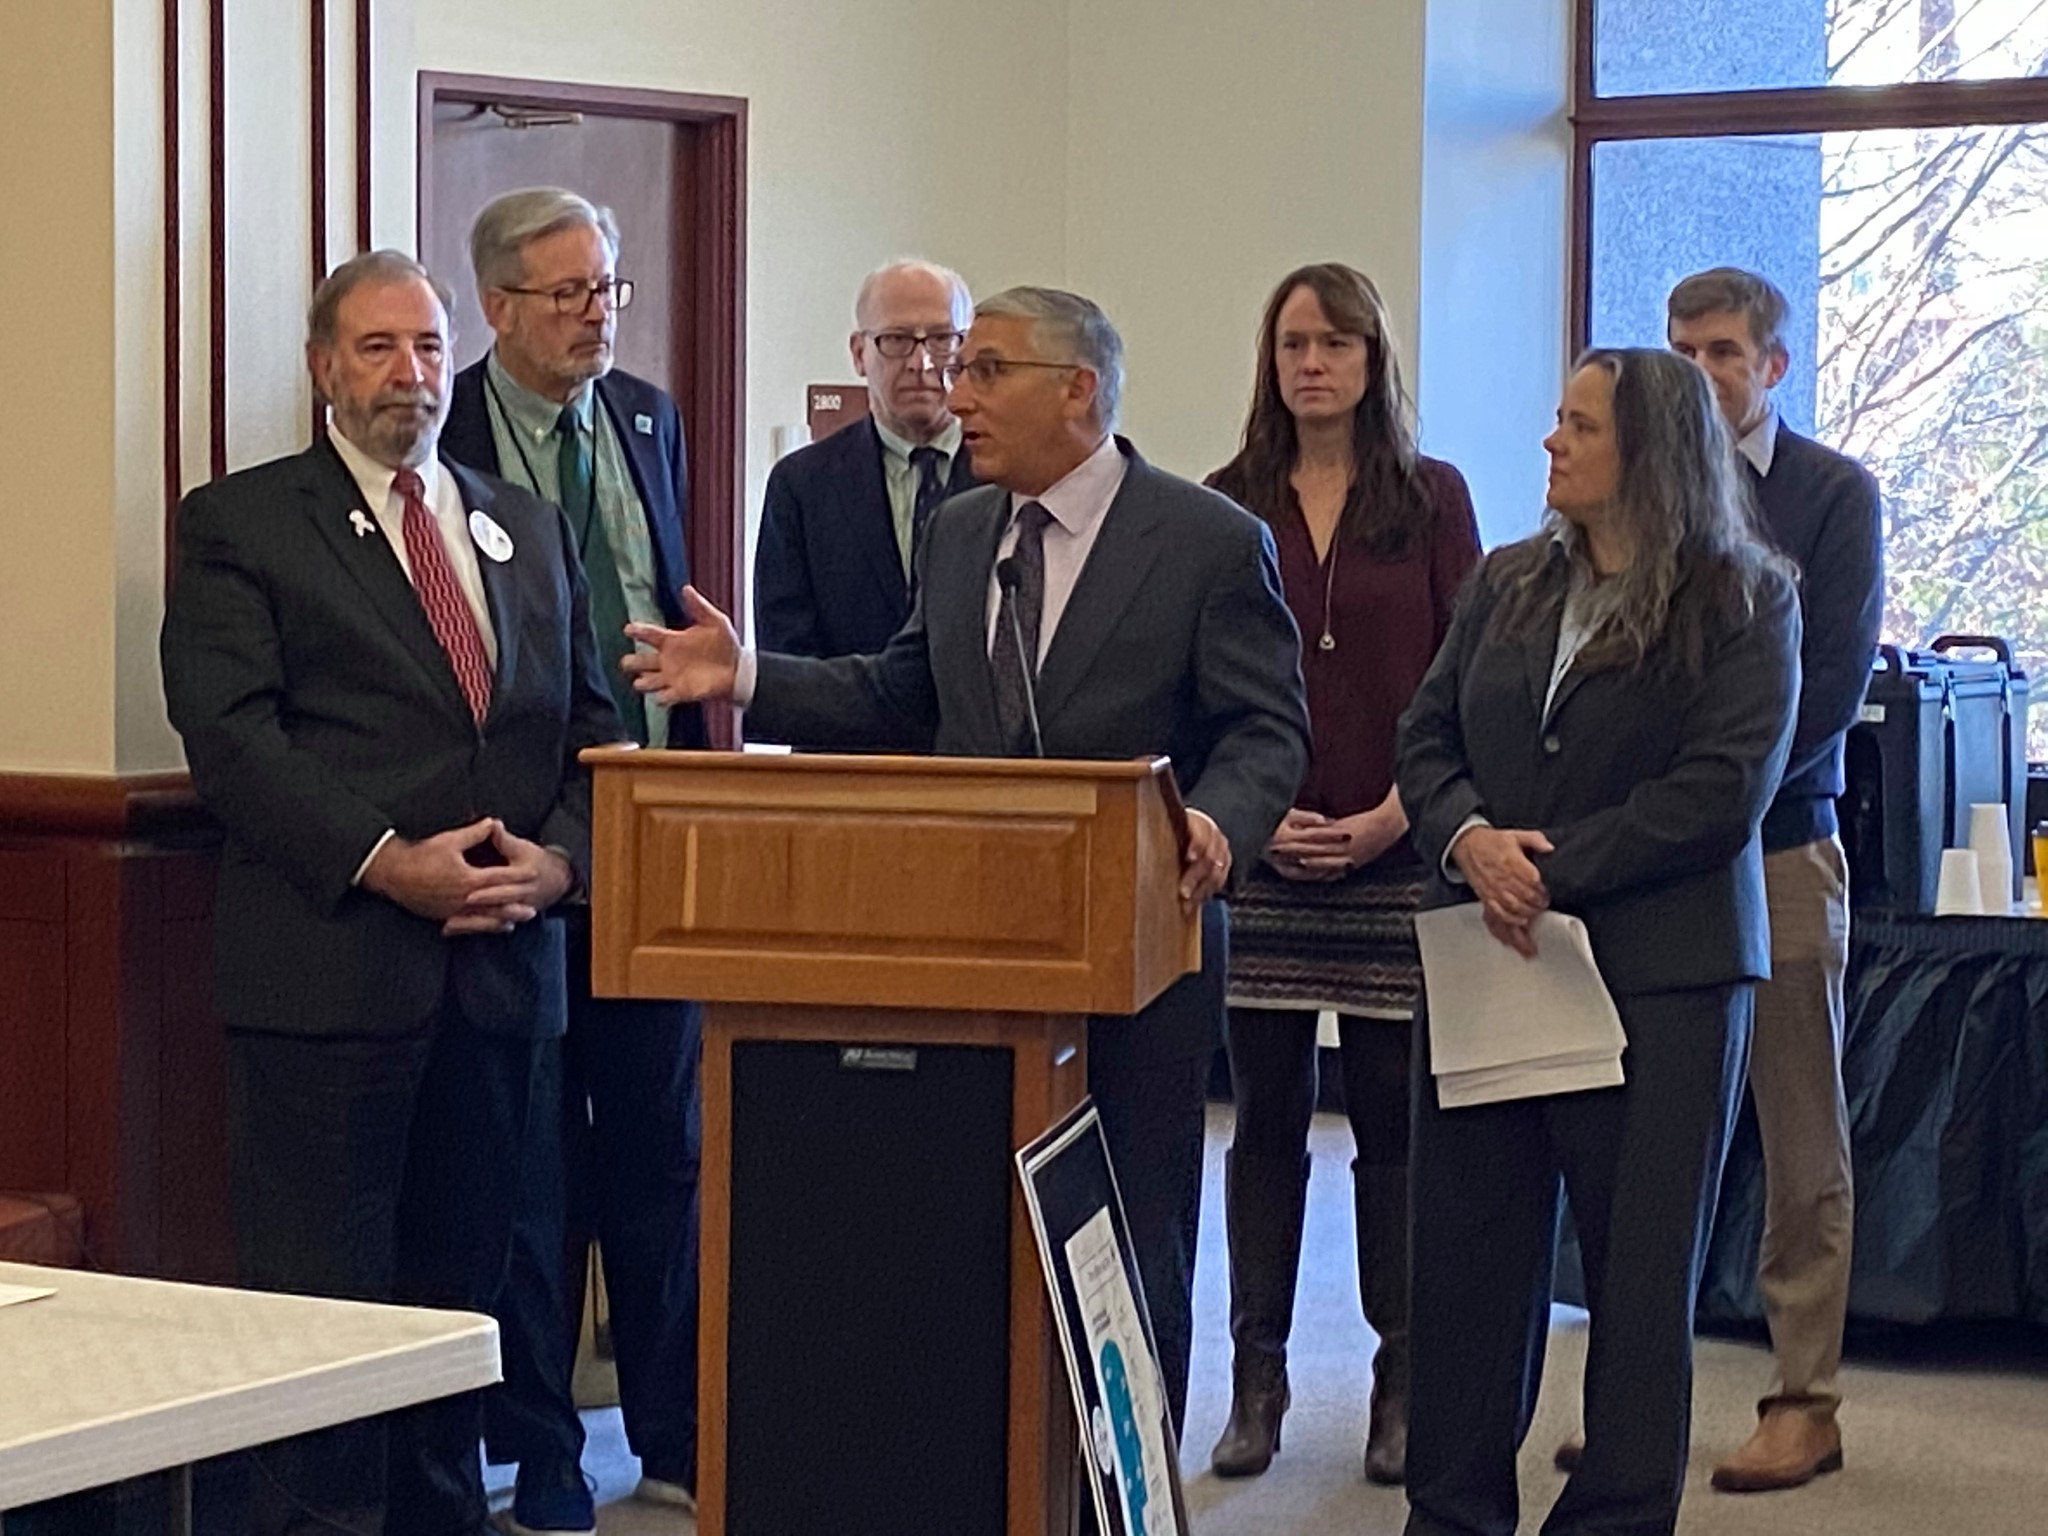
CT Rare Disease Day at the state Capitol in Hartford. Photo courtesy NORD.

The National Organization for Rare Disorders (NORD) and its Rare Action Network (RAN) organized the event for Rare Disease Day, celebrated nationally and in more than 85 countries. Click here for information on your state’s events, RAN and Rare Disease Day.

A disease is defined as rare in the United States if it affects fewer than 200,000 Americans. As many as 7,000 rare diseases exist nationally, affecting 1 in 10 people. Between 25 million and 30 million Americans live with a rare disease, including about 300,000 in Connecticut alone. 

Sofia Sees Hope, based in Ledyard, Conn., gave information to legislators on the Connecticut General Assembly’s Public Health Committee, letting them know that rare disease advocates and those living with a rare disease, such as Leber congenital amaurosis (LCA) and other rare inherited retinal diseases (IRDs), need state and federal support in crafting legislation to help the rare disease community. 

Awareness helps research

We wanted to impress upon the committee that dedicated, grassroots attention and awareness to specific rare diseases generate incredible results in finding cures and treatments. Children living with visual impairment now can regain their vision through a ground-breaking retinal medicine called LUXTURNA. 

The legislators learned that for six years Sofia Sees Hope has been generating awareness, supporting affected families and raising funds to advance research for diagnosis, treatments and cures for blindness caused by LCA and other IRDs, such as retinitis pigmentosa (RP). 

Here’s our legislative statement in its entirety: 

LCA is characterized by severe vision loss at birth. While some children are born with little or no vision, others may have significant vision loss in the first few years of life, stable vision for a period, and then eventually complete vision loss as the retina deteriorates into total blindness.

The optimal window for reversing vision loss is during the early phase of the disease. Creating avenues to affordable treatments and accessibility to resources is imperative and often can be inhibited by insurance regulations and other rules limiting access to help and support patients.

More than 25 genes are associated with LCA and a mutation in just one of these can result in blindness. The rare disease occurs in 1 in 33,000 to 1 in 88,000 people and makes up 5 percent of all retinal dystrophies. Twenty percent of children with visual impairment and attending special schools have LCA; it is the second most common inherited retinal dystrophy after retinitis pigmentosa.

A patient needs a confirmed genetic diagnosis to proceed with appropriate treatment avenues. Sofia Sees Hope has given more than $100,000 to provide families, including those in Connecticut, free access to genetic testing and has directed $275,000 to genetic retinal research. Patients also need support from their lawmakers to ensure they receive the quality of life to which they are deserving.

After decades of research and dedicated investment in studies, scientists created a breakthrough genetic therapy that helps restore vision in patients with one of the genetic mutations causing LCA. The U.S. Food and Drug Administration in 2017 approved this treatment – developed by Spark Therapeutics and called LUXTURNA – which also is the first genetic therapy ever in the United States to treat ANY rare inherited disease.

LCA patients treated with LUXTURNA experienced dramatic changes in their lives with greatly improved or restored vision. Children who are 5, 6, 7 years old and have been treated with LUXTURNA view life in a new light in big and little ways. They now can see rainbows in the sky and stars shining at night.

Our lawmakers need to know that we fully support the principle that all FDA-approved treatments should be made available to all those who will benefit from such treatment, and to reject any proposed requirements restricting access to medications.

Sofia Sees Hope also encourages the Connecticut General Assembly to establish a Rare Disease Advisory Council comprised of patients, patient advocates, doctors, researchers and community members to address the emerging public health priority of rare diseases, including LCA.

More than 20 people – legislators, patient advocates, patients, caregivers, researchers, doctors and businesspeople – spoke during the morning event to a crowd gathered in the second-floor atrium of Connecticut’s Legislative Office Building. 

Jean Kelly, co-founder of Brian’s Hope, spoke on behalf of her son, Brian, and others with Adrenoleukodystrophy (ALD), an x-linked metabolic disorder that destroys myelin, the protective sheath that surrounds the brain’s neurons – the nerve cells that allow us to think and to control our muscles. She and her husband are 24/7 caregivers of Brian who was diagnosed at age 6 and is now 31. Their son can hear but he cannot speak, see or walk. She advocated for more help for parents who must devote their lives to caring for their children. She and her husband advocated for mandatory ALD newborn screening in Connecticut, which was passed into law in 2013.  

Laura Morris from the state’s Office of Health Strategy thanked legislators for the passage 10 years ago of a law requiring health insurance coverage for wound care for people like her daughter, who lived with Epidermolysis Bullosa (EB), a group of rare genetic conditions that result in easy blistering of the skin and mucous membranes. 

A host of other speakers talked about dealing with enormous monthly bills, tangles over insurance coverage and the overwhelming stress on rare disease patients and caregivers.

NORD Director of State Policy Heidi Ross, in a statement from the organization’s President and CEO Peter L. Saltonstall, told the group:

“The purpose of Rare Disease Day is to bring patients and advocates together to articulate with one voice the shared message that millions of people around the world are suffering with unmet medical needs and need help. Our patients need earlier diagnosis; safe, effective treatments; and assured access to medical care and other services …

“There are events like ours today taking place in state capitol buildings across the nation, where elected officials are meeting with patient advocates to better understand what life is like with a rare disease, and how health care decisions they make at the state level – on issues such as newborn screening, medical insurance, cost-sharing and (specially formulated) medical foods – have a major impact” on those living with a rare disease.

Saltonstall’s statement ended with NORD’s credo:

Yes, we are.

Connecticut Rare Disease Day 2020

Rare Disease Day banner with zebra stripes. "Make an Impact on Rare Disease Day! Show Your Stripes."

The end of February signals the time to focus awareness on rare conditions by celebrating Rare Disease Day, a global event addressing the thousands of rare diseases that affect one in every 10 Americans.

Rare medical conditions affect 300,000 people in Connecticut, the home state of Sofia Sees Hope, and 30 million nationwide.

Yes, those are astounding figures that call for astounding action to bring attention to the needs of people living with rare diseases, such as Leber congenital amaurosis (LCA), one of the more than 7,000 rare diseases. The day – and its events that happen nationally and in more than 85 countries – fosters awareness needed to drive research for cures and treatments for rare diseases often overlooked by health-policy decision makers and the medical community.

In Connecticut, people living with rare disorders and their families, along with lawmakers, caregivers, advocates, advocacy organizations, healthcare providers, industry leaders and researchers will gather at the Capitol in Hartford to celebrate Rare Disease Day 2020 by highlighting their concerns and seeking help for solutions from state lawmakers.

Sofia Sees Hope plans to relay rare disease advocacy information, including the importance of genetic testing, to legislators on behalf of people living with LCA and other rare inherited retinal diseases (IRDs).

The public is invited to attend this legislative forum and breakfast at the Legislative Office Building, 100 Capitol Ave., Hartford, from 8 a.m. to 11 a.m., Friday, Feb. 28, the last business day before 2020’s official Rare Disease Day, Saturday, Feb. 29. The event takes place in the building’s second-floor atrium that looks out to the gold-domed Capitol building.

The National Organization for Rare Disorders (NORD) is the U.S. sponsor for Rare Disease Day 2020. The Hartford gathering, as with many others across the nation, is organized by NORD and its Rare Action Network (RAN). The focus for this year’s Rare Disease Day is the impact that rare diseases have on patients, families, caregivers, healthcare providers and local communities.   

For more information about this free event, please contact Lesley Bennett, RAN’s Connecticut Volunteer State Ambassador at Lesley.bennett@rareaction.org or 203-829-7650. Also, here is a link to information on all states regarding RAN and Rare Disease Day: https://rareaction.org/resources-for-advocates/state-profiles/

Here’s a look at the prospective speakers 

Dominic Cotton, a father and advocate for those with rare diseases and brain injuries, will emcee the event that begins after the 8 a.m. sign-in and breakfast.

Heidi Ross, NORD’s Director of State Policy, will offer opening remarks, followed by the legislative welcome by state Rep. Jonathan Steinberg, Co-Chair of the General Assembly’s* Public Health Committee; Rep. Michelle Cook,* member of the Public Health Committee; Sen. Len Fasano, Senate Majority Leader; and Jean Kelly of Brian’s Hope, a non-profit she and her husband, Jack, founded in 2012 for their son, Brian, diagnosed at age 6 with Adrenoleukodystrophy (ALD), an x-linked metabolic disorder. The couple advocated for mandatory ALD newborn screening in Connecticut, which was passed as law in June 2013. 

Dr. Karen Rubin, a pediatric endocrinologist at Connecticut Children’s Medical Center (CCMC), and Adrienne Manning, Connecticut’s Newborn Screening (CT NBS) Program Division Director in the Department of Public Health Laboratory, will address diagnoses and treatments in newborns. State law requires all newborns to be screened for certain genetic and metabolic disorders. The program’s efforts help prevent disability and premature death by ensuring newborns receive the screening and, when needed, evaluation and treatment.

Dr. Rubin and Manning are part of a new partnership between the NBS Program and CCMC called the Connecticut Newborn Diagnosis and Treatment Network.

Silvia Vilarinho, MD, PhD, and Donna Sciacca of the American Liver Foundation are  scheduled to talk about rare liver disorders. Dr. Vilarinho, Assistant Professor of Medicine (Digestive Diseases) and of Pathology at Yale University, is a physician-scientist who uses genetics, genomics and human samples to investigate the molecular basis of various liver diseases of unknown causes. Sciacca is the Community Outreach and Education Manager for the foundation’s Connecticut division.

Dr. Joanna Gell of Jackson Laboratories and CCMC will address germ cell tumors, which can be cancerous or noncancerous growths that form from reproductive cells. Dr. Gell is Assistant Professor in the Department of Pediatrics at the UConn School of Medicine.

Dr. Charles Whitaker, a neurologist who sees patients at the Hospital for Special Care (HSC), will talk about adult neuromuscular disorders.

Laura Morris, mother of a patient with epidermolysis bullosa (EB), a group of rare genetic conditions that result in easy blistering of the skin and mucous membranes, will appear with Rep. Russ Morin, who authored legislation that helped EB patients. Morris is Outreach Coordinator for the state’s Office of Health Strategy.

Dan Donovan, Co-Founder and CEO of rareLife solutions, will address the scarcity of literature on rare diseases.

Father Nikolas Karloutsos will moderate a pediatric panel about the impact of pediatric rare diseases on families. He is a caregiver for his daughter who has a BRAF mutation Rasopathy – probably Cardiofaciocutaneous (CFC) Syndrome – which causes issues with behavioral health and cognition. 

Taking part in the panel will be: 

  • Heather Knapp, caregiver and mother of four, whose youngest was identified at birth through the NBS Program with Phenylketonuria (PKU), an inherited disorder that increases the levels of a substance called phenylalanine in the blood. If untreated, phenylalanine can build up to harmful levels in the body, causing intellectual disability and other serious health problems.
  • Jim Kubicza, who has a son with Angelman syndrome, a complex genetic disorder that primarily affects the nervous system.
  • Information that will be shared on behalf of Marissa B., mother of a child with CDKL5, a neurodevelopmental and epileptic encephalopathy disease characterized by difficult-to-control seizures that begin in infancy, followed by significant delays in many aspects of development. She and her husband provide 24/7 care for their child, who is among those on a years’ long waiting list for a special children’s waiver for Medicaid coverage.  

A panel on the impact on adults with rare diseases will follow, and includes:

  • Carmen Wooster, mother of a daughter with Stiff Person Syndrome (SPS), a rare, progressive syndrome that affects the nervous system, specifically the brain and spinal cord.
  • Beverly St. Onge on Common Variable Immune Deficiency (CVID), one of the most frequently diagnosed primary immunodeficiencies, especially in adults, characterized by low levels of serum immunoglobulins and antibodies, which cause an increased susceptibility to infection.
  • Input on DiGeorge syndrome, a chromosomal disorder that results in poor development of several body systems.
  • The Leeds family regarding hereditary angioedema, a disorder characterized by recurrent episodes of severe swelling.

Paul Pescatello is also set to speak. Pescatello is President and CEO of the New England Biotech Association and chairs Connecticut Business & Industry’s Bioscience Growth Council.

Giving Tuesday: Help Provide Access to Free Genetic Testing

When our daughter Sofia was 2, we knew something was wrong with her vision. By the time she was 5, doctors told us she perhaps had Leber congenital amaurosis (LCA), and the prognosis was grim: total blindness by adulthood. They did say that if we could confirm her condition genetically, there might be a chance for treatment. So for more than seven years, we underwent genetic testing in the hopes of determining which genetic variation was causing her vision loss.

That was more than five years ago. Since then, so much has happened, ranging from our decision in 2013 to create Hope in Focus (formally Sofia Sees Hope) to fund research into treatments, to the fact that in 2019, a child can be diagnosed with a specific type of LCA and receive treatment to restore vision within the same year.

In the time since we learned of Sofia’s diagnosis, 27 different genetic variations have been identified as causing LCA. One – RPE65 – has an FDA-approved treatment that has been performed on dozens of patients with great results. More good news: there are more than 30 clinical trials currently underway for inherited retinal disease, including several promising LCA treatments. But our work is not done. There are 26 LCA genetic variations still awaiting an approved and effective treatment. In the last week alone, we’ve heard from two individuals who have been turned down by insurance for genetic testing to
diagnose their retinal disease.

On this Giving Tuesday (December 3), we are asking for your help in making sure that every single person whose doctor tells them they have an inherited retinal disease has the ability to access a genetic test, easily and at no cost. To date, we have donated $105,000 to make it possible for families to access free genetic testing. That’s more than 100 patients we’ve helped open the door to the next steps in their journey. But there are many more still facing a closed door. We want to open it.

Between now and December 31, every dollar you donate to Sofia Sees Hope will go toward supporting free genetic testing. We thank you in advance for your support.

Rare Disease Advocacy: There’s Power In Numbers

Tell your story. Tell your story again. Then tell it again.  

That’s the beginning of advocacy for rare disease. 

“You have to be assertive and speak up. You don’t have time to waste!” advocate Terri Booker implored her audience at the Sofia Sees Hope second LCA Family Conference.  

Booker, a lawyer and an advocate for people living with Sickle Cell Disease (SCD), spoke as part of a four-member, patient-advocacy panel called “Your Voice Matters!” moderated by Hope in Focus (formally Sofia Sees Hope) Executive Director Annette Tonti.

More than 80 people from 15 states and Mexico attended the July 27 conference in Philadelphia. The event brought together people living with Leber congenital amaurosis (LCA), inherited retinal diseases (IRDs), SCD, which is a group of inherited blood disorders, and Barth Syndrome, a rare genetic disorder that can cause heart failure, muscle weakness and infection.

These diseases are among the 7,000 rare diseases affecting 25 million Americans. 

Kristen Steele hired a lawyer and filed a lawsuit in order to make her career dreams come true. Read her story here.

“Imagine the impact if we all get together and say, ‘We’re here! Twenty-five million people who can vote!’ ” said Booker, who works professionally and personally to help seek justice in the Philadelphia community.

Live long with advocacy

She co-founded the Young Adult Sickle Cell Alliance after her last hospitalization for SCD-related problems in 2012. Doctors with no answers asked what she usually did when symptoms occur, and she replied: “I thought I was here to get help from you.”

SCD is a group of disorders that affects hemoglobin, the molecule in red blood cells that delivers oxygen to cells throughout the body, according to the National Institutes of Health (NIH). People affected have atypical hemoglobin molecules called hemoglobin S, which can distort red blood cells into a sickle or crescent shape.

“I’m talking about it every chance I get,” she said.

If someone notices she’s limping, she tells them she’s got sickle cell, a genetic disorder, “where a whole bunch of cells get together and attack your body, your heart, your eyes, your joints.”

“It’s about accommodating you and whatever it is you’re going through, to make it easier to function.”

Young people with SCD especially need to be empowered to speak up for what they need when they need it and Booker focuses on youth for a big reason: “As a child (with a rare disease), they love you. Once you become an adult, people don’t care about you anymore.”

She encouraged advocating for rare diseases by taking part in events surrounding Rare Disease Day, celebrated annually the last day of February. Rare Disease Week in Washington, D.C., and a multitude of events happen nationally and globally around that time.  

“Help sickle cell patients live long with advocacy,” she said.

The more members of Congress and state legislatures hear stories from rare disease patients and advocates, the more progress will be made toward funding research to find treatments and cures.

“Don’t be afraid,” Booker said. “No one can tell your story but you.”

Panel member Jill Dolgin, PharmD, Head of Patient Advocacy at Applied Genetic Technologies Corp. (AGTC), said rare diseases are small diseases, and suggested a way for people with rare disease to be heard above the noise of research news on more familiar diseases.

“I call it building an orchestra,” – talking about LCA, inherited retinal diseases (IRDs), blindness and genetic testing on the internet, in social media and at conferences – so you’re all “singing the same song” and constantly increasing the volume and frequency of the messages in the song. 

AGTC is developing genetic therapies to treat patients with rare inherited conditions, with its most advanced therapy programs designed to restore visual function and meet the needs of patients with rare blinding conditions, according to the company’s website.

Dr. Dolgin, a healthcare professional with more than 20 years’ global experience in public policy and patient and professional advocacy, said she brings the voice of the patient to small- and medium-sized biotechs to ensure that the needs of the patient are considered and incorporated into every aspect of drug development.

“I’m really helping one patient, one family at a time.” 

Panelist Jamie Ring is Head of Patient Advocacy at Spark Therapeutics, developer of LUXTURNA™, the first approved drug in the United States and Europe to treat an inherited genetic disease and to treat the RPE65 gene which, when mutated, causes one of the more than 25 forms of LCA. She previously worked at Genzyme, a Sanofi company, where she led rare disease patient advocacy and humanitarian programs. 

Ring said her role at Spark Therapeutics is three-fold. 

She serves as a liaison between the company and rare disease communities: “Serving as the voice of the patient inside the walls of our organization.”

She hears directly from the community: “At Spark, it’s really critical for us to understand what the needs of the patients are. You all have a voice at that table.”

Ring also helps connect different rare diseases together to learn how they develop and progress.

LUXTURNA came to fruition after 12 years of research and more that $500 million in investment. The U.S. Food and Drug Administration gave its approval in December 2017. 

“Be involved,” Ring advised the gathering. “Understanding what matters to you, matters to us.”

Panelist Emily Milligan, Executive Director of the Barth Syndrome Foundation, said she is socially and medically committed to serving underserved populations. Before leading the foundation, she launched an $80 million venture fund developing products for Type 1 diabetes and worked with the Juvenile Diabetes Research Foundation overseeing an annual $100 million research portfolio.

Barth Syndrome primarily affects boys and is a multiple-system, complex disorder caused by a chromosomal mutation. It can cause growth delay, impaired lipid metabolism and extreme fatigue. Severe symptoms can result in needing a heart transplant, contracting potentially lethal infections and even death.  

The syndrome, Milligan said, can turn a scraped knee into a trip to the emergency room with sepsis, a life-threatening condition occurring when the body cannot fight infection.

The foundation, she said, became the 14th organization to host a high-level meeting with the FDA, where more than 25 percent of the Barth community, which is about 250 worldwide, told their stories. In research, a second clinical trial is underway toward changing the biology associated with Barth that could mean an improved quality of life for some.

“You have to come together,” Milligan told the group. 

This family conference in Philadelphia has such an enormous impact when people gather together for a common goal, she said. 

“You have no idea the power in numbers. You are a community.”