LCA10 CEP290 Illuminate Clinical Trial Produces No Observed Benefit

ProQR Therapeutics’ clinical trials of sepofarsen to treat a form of Leber congenital amaurosis (LCA) caused by a mutation in the CEP290 gene did not meet its primary endpoint of improving visual acuity.

Illuminate, the name of the clinical study, aimed to explore whether the investigational medicine sepofarsen was effective and safe for people with LCA10 caused by the CEP290 mutation.

The Phase 2/3 clinical trial produced no observed benefit in visual acuity for participants receiving the treatment versus those in the study not receiving the treatment. Visual acuity is a measure of the ability of the eye to distinguish shapes and details of objects at a given distance.

ProQR Founder and Chief Executive Officer Daniel A. de Boer delivered the news recently and characterized the results as disappointing.

“Given the results observed in earlier studies of sepofarsen, the Illuminate trial results are unexpected and disappointing, especially for people living with LCA10,” de Boer said.

“ProQR was founded with the goal of developing RNA therapies  (ribonucleic acid)  for patients with high unmet medical need, and we will continue to advance our robust pipeline of therapies for genetic eye disease. We are deeply grateful to all of the participants, their supporters, and investigators who participated in the Illuminate study.

“Since the results in February, ProQR has been conducting additional analyses of Illuminate and will present these findings at a future scientific conference.”

ProQR works on developing RNA therapies to treat LCA and other inherited rare diseases (IRDs). With DNA being the library of our genes and RNA being a blueprint of that collection, RNA therapies help carry out DNA instructions to create certain proteins critical to a healthy cell.

Analyzing Results for More Answers

Hope in Focus Co-Founder and Board Chair Laura Manfre said we will share updates with the LCA community as we learn more from ProQR.

“The results are not what we had hoped for from ProQR on the LCA10 Phase 2/3 trial, but we are resolved to keep hope in focus, knowing that the brilliant minds that got us this far are not giving up.

“It’s important to remember when there are setbacks that only 10 years ago, all we had was hope. Most of the community couldn’t even get a genetic diagnosis, let alone hope for treatment,” she said. “We’ve come a long way in a short period of time, and we’re going to keep advancing.”

Ben Yerxa, PhD, Chief Executive Officer of the Foundation Fighting Blindness, said his organization will work with ProQR to learn more about the data.

“This was not the outcome we had hoped for, and we share in the disappointment many are feeling in the community,” Yerxa said.

“We will continue to work alongside ProQR to learn more from the ongoing analyses and as they work to advance RNA therapies to potentially help children, adults, and families who are affected by rare genetic eye diseases.”

In delivering the results, the biotechnology company gave background about LCA, the most common genetic cause of childhood blindness, affecting about 15,000 people in the Western world. One federally approved gene therapy treatment, LUXTURNA®, exists for people with LCA2 (RPE65), one of the more than 27 forms of LCA.

The rare retinal disease usually appears in the first year of life and is characterized by progressive loss of vision. Other symptoms can include rapid eye movement, known as nystagmus, eye-poking, night blindness, and sensitivity to light, known as photophobia. Depending on the mutation, complete loss of vision can occur during early childhood.

Specifics of the LCA10 CEP290 Trial

Illuminate enrolled 36 participants, aged 8 years or older with genetically confirmed LCA10 due to the c.2991+1655A>G (p.Cys998X) mutation in the CEP290 gene.

The study was a randomized, sham-controlled clinical trial that took place in three randomized groups at 14 sites in nine countries.

The first group received a target dose of sepofarsen by intravitreal injection (IVT), the second received a low dose via IVT, and the third underwent a sham procedure that mimicked an injection with no medicine or injection given.

Bart P. Leroy, MD, PhD, one of the study’s key investigators and Director of the Ophthalmic Genetics and Retinal Degenerations clinics in the Division of Ophthalmology and Center for Cellular and Molecular Therapeutics at The Children’s Hospital of Philadelphia (CHOP), said work will continue toward finding therapies.

“LCA10 is devastating, and with no approved therapies, very difficult to treat retinal disease resulting in blindness,” said Dr. Leroy, who also is head of the Ophthalmology Department at Ghent University Hospital and Professor of Ophthalmology and Ophthalmic Genetics at Ghent University in Belgium.

“We will continue to work with ProQR to understand the data as they work for advance therapies for individuals with inherited retinal diseases.”

Andy Bolan, ProQR Director of Medical Affairs, said the team extends its thanks to the study participants, their supporters, the investigators, and their staff for support in developing the trial. He said the biotech remains committed to making a significant and positive impact on the lives of those affected by genetic conditions.

For quarterly news and future study participation opportunities, you can sign up for the ProQR Eye Connect Newsletter or follow them on social media.

People also can contact ProQR with any questions at patientinfo@proqr.com.
Let’s Chat About … Why Natural History and Patient Outcome Studies Matter

Amid the intricacies of researching treatments and cures for rare diseases, such as Leber congenital amaurosis (LCA) and other inherited retinal diseases (IRDs), the patient remains the major focus.

Researcher Jonathan Stokes says he takes a holistic approach in developing and evaluating patient-health outcomes in clinical trials that include focusing on signs and symptoms of disease, health-related quality of life for patients, understanding unmet needs, and exploring the burden of disease.

“Patient voices matter,” he said.

Stokes is Director of Patient-Centered Outcomes Research for AbbVie, a Chicago-based biopharmaceutical company. He holds a Master’s in Business Administration from Northeastern University and has a devoted interest in understanding and bringing to light the patient voice and perspective, with more than 16 years of research study design and implementation experience.

He primarily works in health-outcomes research, specifically developing and evaluating clinical outcomes of assessments (COAs) used in clinical trials to substantiate treatment benefit.

Sofia Sees Hope featured Stokes in an April 19 webinar episode of “Let’s Chat About…Why natural history and patient outcome studies are important.” Elissa Bass, Director of Marketing and Communications for Sofia Sees Hope, moderated the session that is part of the organization’s free, monthly webseries. 

We developed the series with the LCA and IRD communities in mind but invite all members of our community, including those in research, industry, and the regulatory communities to join any of the sessions as we look ahead to a common goal of advancing treatments for rare retinal disease.

Objective Versus Subjective Gauges

Patient-centered outcomes are evaluations of a patient’s health status and provide valuable information on how patients feel and function.

Vision measured by navigating a maze renders a tangible, objective result, as do blood tests that reveal cell counts or measures that evaluate tumor size in oncology. 

Certain conditions, or aspects of a given condition, express themselves in ways only known to the patient, such as fatigue with mitochondrial diseases or pain with migraines. These subjective experiences can be assessed in studies and through outcome measures, such as daily diaries or questionnaires. These assessments involve years of qualitative and quantitative research, and they are designed after consulting with patients, their caregivers, doctors, researchers, and literature. 

This information literally is the patient’s voice.

Stokes says, “For me, the question is always the ‘So what?’ What does that mean to the patient?’ ”

He takes an inclusive approach to developing measurements for patient outcomes, considering quality-of-life elements – the effects of a disease emotionally, socially, physically, and in daily activities – to produce valid, reliable results.

“Even if you are not taking part in a clinical trial, these are all steps that need to be done to make sure we’re doing it the right way.”

The information, in turn, is shared back with the patient community. 

“These kinds of things matter to people,” he said. “Their feelings are not just idiosyncratic to themselves.”

This research is critical to the drug approval process and important to all stakeholders – patients, care givers, researchers, regulators, and drug developers – to create what Stokes calls a true partnership.

Health-outcomes research plays an important role in the U.S. Food and Drug Administration’s Center for Drug Evaluation and Research that specifically targets patient involvement through its Patient-Focused Drug Development (PFDD) program.

“PFDD is a systematic approach to help ensure that patients’ experiences, perspectives, needs, and priorities are captured and meaningfully incorporated into drug development and evaluations,” according to the FDA.

The goal is to better incorporate the patient’s voice in drug development and evaluation, which ultimately results in an FDA-published document called “Voice of the Patient.”

Natural History studies also are important to patient outcomes because they reveal the patient experience over time as researchers observe features in the absence of any treatment. These data give knowledge and an independent understanding of the disease, while establishing an essential foundation for building drug development programs. A Natural History study also can be used as a control arm serving as a placebo in a clinical trial.

The studies track the course of a patient’s disease, identifying demographic, genetic, environmental, and other variables that shape the drug development process. They give scientists and researchers a better estimate of the prevalence of the disease, help identify biomarkers, affect clinical outcome assessments, and determine the feasibility of established assessments for clinical trials. 

More than ever, Stokes said, all stakeholders in the drug development process come together and work toward a solution best for the patient.

“Patients are at the center of everything we do.”

‘Let’s Chat About …’ Webinar Offers LCA Overview and Updates on Clinical Trials

In the debut of Hope in Focus (formally Sofia Sees Hope) ‘Let’s Chat About …’ monthly webinar series, Ben Shaberman of the Foundation Fighting Blindness, provided his Zoom audience with a plethora of information about Leber congenital amaurosis (LCA), highlighting some of the more than 40 clinical trials underway to find treatments and cures for LCA and other rare inherited retinal diseases (IRDs) and giving updates on promising preclinical research. 

The recorded webinar aired 1 p.m. Wednesday, Jan. 27, 2021, and can be seen here. Elissa Bass, our marketing and communications director, moderated the session.

Shaberman, Senior Director, Scientific Outreach & Community Engagement, stumbled across a science writing position at the Foundation Fighting Blindness 16 years ago without a clue about retinas or blindness. He called his move to the Foundation serendipitous. He knew he made the right choice after hearing retinal researcher Dean Bok, PhD, tell attendees at a 2005 Foundation conference how he was drawn to the field by the seduction of the retina’s myriad complexities and inner workings.

Shaberman, too, felt pulled by the intriguing science of the retina.

As such, so are the 27 forms of LCA that cause varying kinds of visual impairment within each gene mutation and within each affected person. An estimated 8,000 people in the United States have LCA.

The path of retinal research

Shaberman took his audience from the beginnings of identifying the RPE65 gene in 1993 and learning shortly thereafter it could lead to LCA, to using mice models and later studying Briard dogs that had the same gene mutation that caused LCA in humans. A clinical trial at Children’s Hospital of Philadelphia led to the 2017 FDA approval of the breakthrough gene therapy LUXTURNA®, developed by Spark Therapeutics. The drug successfully improved the vision of many of the LCA2-RPE65 patients who received the treatment through subretinal injections.

When children receive an LCA diagnosis, their families should find a good retinal specialist, get regular exams, and ultimately get a confirmed genetic diagnosis to be on the path to more specific information and research into that form of LCA, Shaberman said.

Families also should register with the Foundation’s My Retina Tracker®, a free and secure online registry that facilitates getting a confirmed genetic diagnosis by making registrants eligible for free genetic testing.

The registry becomes your personal retinal health record, updated by you. It employs state-of-the-art database technology to protect privacy and adheres to the highest standards of confidentiality and ethics. 

It also notifies registrants of clinical trials and gives researchers access to their disease data – not their personal information – to advance research and therapy development associated with LCA and IRDs. 

Reading research publications and attending events sponsored by the Foundation and by Sofia Sees Hope also provide opportunities for families to interact and learn the latest research. Shaberman and Bass encouraged people affected by LCA and their families to contact them, respectively, through the Foundation’s website and/or the Sofia Sees Hope website for specific information on clinical trials or other questions and concerns about living with LCA. 

“Yes, it’s work,” Shaberman said. “You have to be your own advocate and your own child’s advocate, but more and more information is becoming available, and that’s the good news.”

Shaberman also reviewed some of the more than 40 retinal clinical trials in the pipeline for LCA and other IRDs:

Join us Feb. 16

February’s “Lets Chat About …” webinar airs at 3 p.m. ET, Tuesday, Feb. 16. Our guest will be Wiley A. Chambers, MD, Supervisory Medical Officer for the Office of New Drugs, Center for Drug Evaluation and Research at the U.S. Food and Drug Administration. Register here.

ProQR Completes Enrollment for Next Phase of RNA Therapy for LCA10-CEP290

ProQR Therapeutics reached an important milestone by completing enrollment in the next pivotal phase of clinical trials of sepofarsen, a developing RNA therapy for treating LCA10, a form of Leber congenital amaurosis (LCA) with a mutation in the CEP290 gene

The biotechnology company based in Boston and in Leiden, The Netherlands, announced earlier this month that it finished enrollment in its Phase 2/3 Illuminate study of sepofarsen for treatment of LCA10 due to the p.Cys998X mutation in the CEP290 gene. 

LCA10 is a severe retinal dystrophy, causing blindness or severe visual impairment at birth or during the first months of life. The mutation affects about 2,000 people in the Western world

Sepofarsen is an antisense oligonucleotide (AON)  that works like “genetic tape” to fix the mutation. The therapy is unlike gene replacement therapies in which whole genes are delivered to replace defective copies. 

The drug aims to repair the genetic defect that causes the disease in the ribonucleic acid (RNA). The mutation leads to an aberrant splicing of a person’s messenger RNA (mRNA) and leads to a non-functional CEP290 protein. Sepofarsen is designed to enable normal splicing, resulting in subsequent production of functional CEP290 protein. 

Recruiting patients for clinical trials represents one of the biggest challenges in getting studies underway. The COVID-19 pandemic and concerns about spreading the virus presented an unprecedented challenge in and of itself. Pharmaceutical companies also have steered their focus from studies across the board as they raced toward developing and rolling out worldwide virus vaccines. 

Finding the required 33 LCA10 individuals for the Illuminate trial in a subset of a people with a rare condition was exceptionally challenging, and ProQR exceeded that requirement by enrolling 36 participants.

During the Phase 2/3 study, the 36 patients, ages 8 and older, are set to receive sepofarsen either in a dose expected to be used once the drug gains approval or a lower dose or a placebo. The 12-month clinical trial is intended to support an application for marketing approval of sepofarsen. The study is being conducted in the United States, Canada, Brazil, The Netherlands, Belgium, France, Italy, Germany, and the United Kingdom.

The drug is administered through intravitreal injections in the eye. It is also a platform for use as transformative therapies for treating Usher syndrome and retinitis pigmentosa (RP)

Top-line results in early 2020 from the Phase 1/2 clinical trial of sepofarsen in 11 children and adults revealed that 60 percent of patients had improvements in visual acuity and navigating a mobility course. The study, carried out in the United States and Belgium, also netted a super responder, a person who responded particularly well to the treatment. 

Laura Manfre, chair and co-founder of Hope in Focus (formally Sofia Sees Hope), said that as the parent of a child with an LCA diagnosis, she was told there was nothing that could be done and that her family needed to accept that their daughter would one day be blind.

“Now, in early clinical testing we have seen the potential for sepofarsen to make a significant difference for patients with LCA10 due to a mutation in the CEP290 gene,” she said. “We see hope for individuals living with this disease. We look forward to learning about the results of the Illuminate trial and continuing to work with ProQR as they advance their pipeline of RNA therapies to potentially help children, adults, and families who are affected by blindness caused by LCA and other rare inherited retinal diseases.”

Aniz Girach, MD, ProQR’s chief medical officer, said in a statement that the company was pleased to have completed enrollment of the Illuminate trial of sepofarsen.

“This marks an important milestone for ProQR, as well as for the LCA10 and broader inherited retinal disease community,” Girach said. “In surpassing our enrollment target, we were able to accommodate the broad interest to participate in the trial. This speaks to the fact that there are currently no approved treatments for patients with LCA10.

“If approved, sepofarsen has the potential to be the first therapy to address this high unmet medical need for patients who would otherwise face blindness, he said. 

“We are grateful to those who have supported our efforts in bringing this trial forward, including our investigators, patients, and caregivers. We look forward to sharing the top-line results in the first half of 2022.”

Reflecting on the Trajectory of IRD Research

When I joined the Foundation Fighting Blindness as a science writer in 2004, I really didn’t know what I was getting into. I knew nothing about the retina, let alone the complex and diverse world of rare inherited retinal diseases (IRDs) that includes Leber congenital amaurosis (LCA). But the research for treatments was cutting-edge and compelling, so I was excited to dive in and learn.

My early assignments were writing about laboratory studies coming out of academic labs. There were virtually no companies in the IRD space and only one or two clinical trials underway for emerging therapies. But there were a lot of studies of genetically engineered mice and rat models of IRDs for gaining a better understanding of disease pathways and testing potential treatments.

Truth be told, I often wondered if and when rodent-tested therapies were really going to make it into human studies and out to the people losing vision. But the scientists conducting the research were mind-blowingly smart and innovative, so I figured they knew what the heck they were doing. With a master’s degree in poetry, who was I to judge?

Fast forward about four years: I was in my hotel room in Fort Lauderdale – there for the annual Association for Research in Vision and Ophthalmology conference – when my manager called and told me three research groups just reported vision improvements in young adults treated with RPE65 gene therapies in Phase 1/2 clinical trials. That was the breakthrough we’d all been waiting for.

People, rather than animals, with severe vision loss were now seeing significantly better. It was the first time an IRD treatment had worked in humans. I will never forget the headline for the article I immediately wrote: “Now They See.” (Note: One of those RPE65 gene therapies later became LUXTURNA®, the first FDA-approved treatment for the eye or an inherited condition.)

After many years of painstaking work, our hope for treatments and cures had finally begun evolving into promise.

There have been several other aha! moments in the ensuing years, but I distinctly recall cathartic encounters at the 2019 American Society of Retinal Specialists in Chicago. As I perused the snack table during breaks (the accomplished snacking professional that I am), several representatives from biotechs developing IRD therapies – companies I’d never even heard of – came up to introduce themselves to me and tell me about their emerging IRD treatments. They didn’t know me or my role, nor had I previously known them; they were just eager to connect with someone from the Foundation Fighting Blindness to get on our radar screen.

I realized then I couldn’t keep track of all the companies (dozens) focused on IRDs and clinical trials (40-plus) underway for potential IRD treatments. But being overwhelmed felt incredibly good, and it meant more good news likely was on the horizon for saving and restoring vision.

While mouse studies are as critical as ever, I can’t remember the last time I wrote an article about one. That’s because most of my writing is now dedicated to reporting on advances, including encouraging vision improvements, being made in human studies.

Make no mistake: Much more work needs to be done before we eradicate the myriad IRDs affecting millions of people across the globe. And, of course, we cannot get more therapies across the finish line fast enough. But when I look at how incredibly far we’ve come since those early days of mice and rats, I have no doubt we are well on our way to breaking many more ribbons soon.

Nearly Three Years Later, LUXTURNA®™ Treatments Continue to See Success

Since its launch in March 2018, breakthrough gene therapy LUXTURNA®™ continues to be successful in helping improve vision in people with inherited retinal disease due to mutations in both copies of the RPE65 gene and viable retinal cells as determined by a healthcare professional. The therapy treats LCA2, known as LCA/RPE65, one of more than 25 forms of Leber congenital amaurosis

The drug – developed by Spark Therapeutics and a team of retinal research superstars that included Dr. Katherine A. High  and Dr. Jean Bennett – came to fruition after 12 years of research and millions of dollars in investment. 

Spark Therapeutics could not comment on the number of people who have received the gene therapy, but spokesman Kevin Giordano recently said the company has shipped well over 200 vials of the therapy since the U.S. Food & Drug Administration approval in December 2017. One vial of the drug treats one eye.

Trained retinal surgeons at one of the 10 eligible treatment centers in the United States deliver the gene therapy to the back of the eye by subretinal injection using a needle the size of an eyelash; about a week or so later, the patient undergoes the procedure in the other eye.

Read: 10 US Centers Offering Treatment

“Spark Therapeutics is thrilled patients continue to benefit from LUXTURNA (voretigene neparvovec-ryzl),” Giordano, Spark’s External and Product Communications Lead, said. “A gene therapy is a major milestone, not only for those of us at Spark, but also for the patient community and broader gene therapy field.”

The cost of the drug – $425,000 for each eye – initially caused anxiety among patient families, but Giordano had good news about insurance coverage.

“Payer coverage has exceeded expectations, and to our knowledge no treatment-eligible patient has been denied treatment due to their insurance coverage,” he said.

LUXTURNA also is being used beyond this country through license and supply agreements with Novartis, which has the rights to commercialize the drug in Europe and all other markets outside the United States. Spark manufactures and supplies the gene therapy to Novartis, according to Giordano.

Also, results from ongoing studies continue to support the drug’s safety profile and efficacy.

“In 2019, Spark presented four-year durability data from the LUXTURNA Phase 3 clinical trial,” Giordano said. 

Read: LUXTURNA safety information

For privacy reasons, Spark Therapeutics cannot share patient results.

But we can – at least a couple of them because they came from patients and families sharing their stories with us.

For Hannah, ‘Hope Realized’

Hannah Reif, daughter of Amy and Chris Reif of Maple Glen, PA, underwent LUXTURNA gene therapy in July 2018. Dr. Albert M. Maguire, who is married to researcher Dr. Bennett, performed Hannah’s surgery at Children’s Hospital of Philadelphia (CHOP).  

Hannah with a big smile, wearing a light blue green coat and holding a drink
Hannah Reif

“We closely watched the clinical trials and the FDA approval process for seven years, starting when Hannah was diagnosed with LCA/RPE65 at just a few months of age,” Amy said. “Seven years of hope. 

“Two years out from Hannah’s surgery, I can say we feel grateful and fortunate that she was treated with LUXTURNA. No regrets. LUXTURNA was hope realized. It delivered what it promised.”

She said what that has meant for Hannah has been nothing short of life changing.

“It has given her more independence, which has been wonderful for her self-confidence. It has given her the ability to see what she couldn’t before.”

Since the surgery, Hannah’s vision in dim lighting and her visual acuity improved. She is now 9 and just finished third grade. 

Sometimes, her mom said, it’s the little things that are the most amazing.

“A year after the surgery, she was about to eat hot oatmeal and said, ‘Hey, I see steam. Hey, I can see that,’ ” Amy said.

“There are still things that pop up that she’s seen for the first time, like when she said, ‘Mom, did you know there’s a pattern on this toy?’ It’s fun to see her discovering.”

Amy said she and her family will be forever grateful to Dr. Bennett.

“There has been a lot of talk about heroes recently and Dr. Bennett is our hero. We are grateful for this groundbreaking treatment that she developed, that has been life-changing, not only for our daughter, but also for the sons and daughters and loved ones of so many others as well.”

‘His Vision Changed Everything’

Creed Pettit, one of the first recipients LUXTURNA, received the gene therapy at age 9 in March 2018. Dr. Audina M. Berracol  performed the surgery at Bascom Palmer Eye Institute in Miami, Fla.

His mom, Sarah St. Pierre Schroeder, told us that her now-11-year-old is doing amazing and only occasionally has issues with dim lights, “but nothing like before.”

Their days in Mount Dora, Fla., have changed in a major way.

“He still starts his day with his trusty smoothie and waffle, but after that, Creed wants to create new pranks (today was putting ice in the tub). He said it was nice and warm so I could soak my feet.”

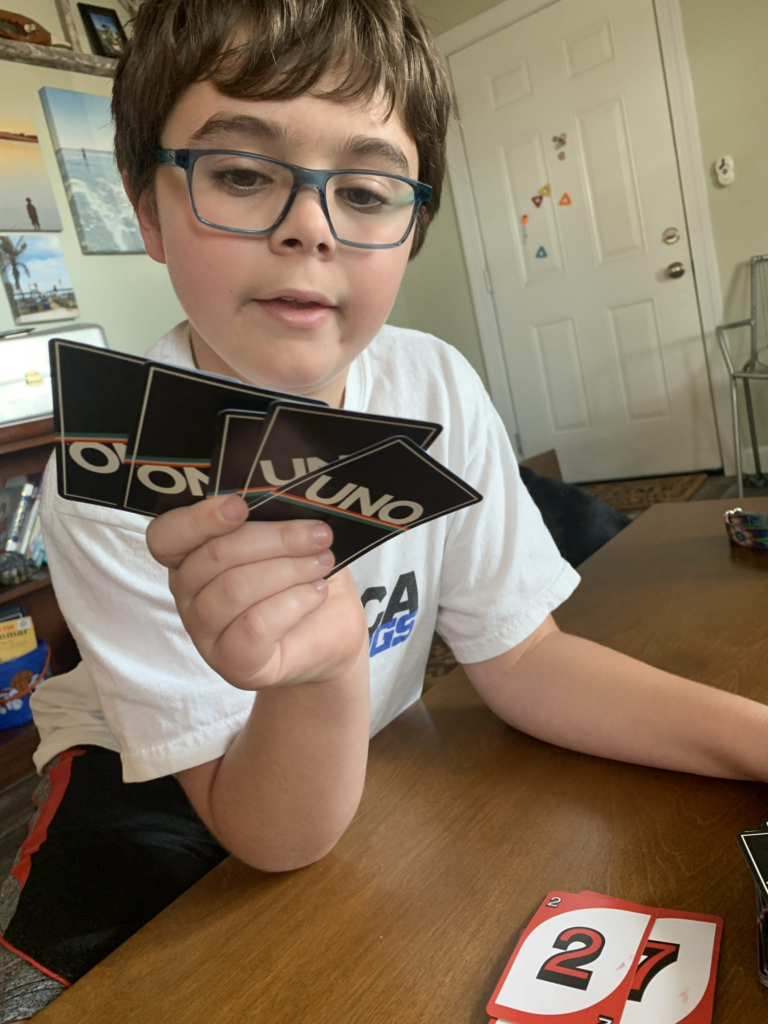
Creed in blue glasses, holding his Uno Cards
Creed Pettit

Creed now loves to play board games – Trouble, Sorry, Battleship and Uno. 

“His vision has changed everything. He can manipulate small objects, he is using pointer fingers, and loves trying to roll his eyes in the mirror.”

Creed still likes to draw, and he loves riding and popping wheelies on his bike named Carlitos. He also likes to talk.

“Talking more is an understatement! Sunup to sundown, he is talking. He has also learned how fun it is to use his imagination, something he never did before. He creates awesome stories during this time.  He is so much more independent; I have to remind myself of that often.”

At first, she and her husband, Chad, could see that Creed’s vision improved some, but once he became confident about his gene replacement, they noticed him finding toys and games.

“He was suddenly enjoying things he didn’t before. He now expresses when he can’t see. Yesterday it rained all day, I kept waiting for him to tell me it was too dark inside, but he was fine. He just started doing staring contests. I love looking at his eyes during the contest.”

Sarah said she is incredibly grateful.

“Creed’s surgery is something we still thank God, St. Raphael, St. Lucy and Sister Miriam Teresa, for every night.” 

Like many kids across the country, Creed finished school at home because of the coronavirus pandemic. He graduated from fifth grade and thought because he finished school at home, he wouldn’t have to wear the graduation cap.

“But I was not going to let that slide,” his mom said. “I made one and took pictures.”

Throughout his journey, though, Creed felt anxious, something Sarah had learned that might happen when, before Creed’s surgery, she talked with Tami Morehouse, a LUXTURNA clinical trial participant at age 44.

Tami, a Sofia Sees Hope Ambassador, tries to calm fears and advises potential gene therapy patients and their parents that even though undergoing the surgery has the potential to do such good by improving vision, they should think about their expectations, especially with children.

“We are comfortable in our own zone; give us a little bit of change and it can throw us off,” Tami said.

Sarah understands. 

“He started to have a lot of anxiety. He had a hard time sleeping. I feel everything changed so fast that he was overwhelmed, but we have worked hard on getting past that.”

Sarah and her family are in a great place now. But after they were home from the hospital in Miami and settled into their routine, she said she became very emotional.

“All the tears I had held in for nine years started to come out. I felt I no longer had a purpose; I was so used to staying busy. I did not know who I was supposed to be.” 

She got some help and realized she still was needed because people need her help learning about and understanding this groundbreaking gene therapy.

“I still find myself shocked over how this has changed Creed’s life and so many other lives.”

Clinical Trials and Emerging Research Show Promise for LCA Treatments

Forty clinical trials and a lot of pre-clinical research into LCA treatments show promising pathways to discovering the next LUXTURNA®, according to Shannon Boye, PhD, the opening speaker for the Virtual VISIONS 2020 conference, presented earlier this summer by the Foundation Fighting Blindness

The breakthrough drug developed by Spark Therapeutics marked a milestone in the history of genetic research as the first gene therapy in the United States for any inherited disease and as the first to treat one of the more than 25 forms of Leber congenital amaurosis (LCA).

Shannon Boye, PhD in a lab coat
Shannon Boye, PhD

Boye, along with Foundation Chairman of the Board David Brint and Foundation Chief Executive Officer Benjamin Yerxa  PhD, kicked off the three-day, first-time virtual conference, the Foundation’s major annual gathering. Rather than convening in person, the event’s speakers, exhibitors and more than 1,600 attendees participated through an online app, due to concerns surrounding the coronavirus pandemic.

Brint said that the 40 clinical trials and more emerging treatments for various IRDs span the disease profile.

“No matter what your disease is, these hopefully will be able to restore vision,” Brint said. “In the next 10 years, we have an opportunity to bring many more vision-saving treatments into and through the pipeline and across the finish line.”

Yerxa said the topic of genetic therapies would be good to lead off with because of the sheer variety of innovative programs and approaches to each therapeutic challenge.

“There’s essentially a revolution happening right now in personalized medicine and genetic therapies in general,” Yerxa said.

Boye, an assistant professor in the Department of Ophthalmology at the University of Florida, addressed the audience in the beginning session called: “Mission Possible! What’s Next?”

She discussed three major strategies in treating LCA and other IRDs: Gene supplementation or gene replacement therapyRNA therapeutics and gene editing

Boye set up an analogy to better understand the complexities of these strategies, saying we all have little letters in our bodies called DNA. Subunits of those letters – that DNA – are genes. RNA carry the instructions from DNA for making proteins, the building blocks of life.

“They act alone or in concert with a bunch of other proteins to perform essential functions.”

Continuing her letters analogy, Boye said, imagine a friend texts you: ‘Please take out the dog.’ You get that message and perform that function because letters combined correctly to tell you to take the dog out. 

If only the word ‘Please’ appears on your phone screen, you don’t take the dog out.

Or, if the ‘d’ is pushed and an ‘l’ comes out, sending the message, ‘Please take the log out,’ “you then have a mess to clean up,” she quipped.  

In the first strategy of gene supplementation or gene replacement, the right protein needs to be expressed in the patient’s retina. 

The letters need to be correctly sequenced to generate a coherent message, in this case, telling a protein to perform an important function. Any break in that cascade of events can cause visual impairment. 

The gene replacement therapy LUXTURNA is a human-engineered virus containing copies of the corrective gene that doctors deliver through a subretinal injection so the cells can make the originally missing protein.

“You deliver the right letters that make the right message and the right protein,” she said. “That’s a pretty simple concept. That’s LUXTURNA.”

Developed to improve vision in people with LCA2* caused by a mutation in the RPE65 gene, LUXTURNA received Food and Drug Administration approval for  use in humans in December 2017. 

One area of Boye’s research as Associate Division Chief of Cellular and Molecular Therapeutics is entering into a Phase 1/2 clinical trial, applying the same premise for mutations in the GUCY2D gene that causes LCA1

“It’s early,” she said. “But this is an example of another perhaps next LUXTURNA being right around the corner.”

She cited similar research moving forward on other IRDs, including Retinitis Pigmentosa (RP)Choroideremia, and Bardet Biedl Syndrome (BBS)

The second strategy is a form of RNA therapeutics that uses antisense oligonucleotides (AONs) – short, single-stranded DNA  molecules that interact with messenger RNA to correct translation of a targeted gene. Think of an AON as an autocorrect feature that binds to the ‘l’ in log and changes to a ‘d’ for dog.

Promising pre-clinical work now in Phase 2/3 for CEP290 or LCA10 also is coming out of Rob Collin’s research group in The Netherlands, Boye said.

Another AON program underway addresses a form of Usher Syndrome

The third strategy – the newest and most exciting – is gene editing. A guide RNA is used to drag a special enzyme to a region in the DNA that contains the mutation, and the enzyme cuts the DNA, like molecular scissors. 

Researchers are exploring a host of gene editing variations, including cutting out a specific area of DNA and replacing it with the right letters to make a coherent message. The lab work has created paths to address a range of IRDs, including CEP290,  Usher Syndrome,  RP, Stargardt Disease and Choroidermia.

“There’s an absolute exponential increase in the therapies that are being developed,” she said.

These strategies are not limited to the disease conditions under discussion and can be more widely applied to a number of genes and conditions.

Addressing those who do not have RPE65 or LCA2 for which a treatment exists, Boye said, with all of this research in progress, “that one day, there’s going to be a LUXTURNA for your inherited retinal disease, too.” 

Living with LCA: Finding Her ‘Light’ In the Kitchen

A recipe for addiction recovery transformed Orly Shamir’s life, and now it’s about to change her future.

Orly, who’s name in Hebrew means “My Light,” lives with LCA4, a form of Leber congenital amaurosis caused by a mutation in her AIPL1 gene. 

The 52-year-old Canadian, newly transplanted to southeastern Florida, takes her Hebrew name literally.

“I am light, and I am vision,” she said. “I want to offer everything I have for others to realize their perfect light and vision is possible.”

As a child she had enough vision to read large print, but in her 20s, Orly’s sight deteriorated to minimal light perception and shadows. In 2014, she was part of a clinical trial in Canada for the Argus Retinal Prosthesis System (Argus II). The Argus II, known as the bionic eye, stimulates the eye with electrodes to transmit visual information captured by a video camera to the patient’s brain. You can read about her experience in the trial here.

“Still, I have true 20/20 vision,” she said. “My blindness forces a mindful clarity through all my other senses and that enriches everything from my cooking and healing, to my service to others.”

Along her journey through the darkness of an opiate addiction, she rediscovered her mother’s traditional Moroccan fish dish, served as part of each Friday’s sabbath dinner. 

Little did she know the importance this recipe would have to her survival.

From 1999, after having her third child, until 2012, she said, “My opiate addiction took a huge chunk of my life away because it

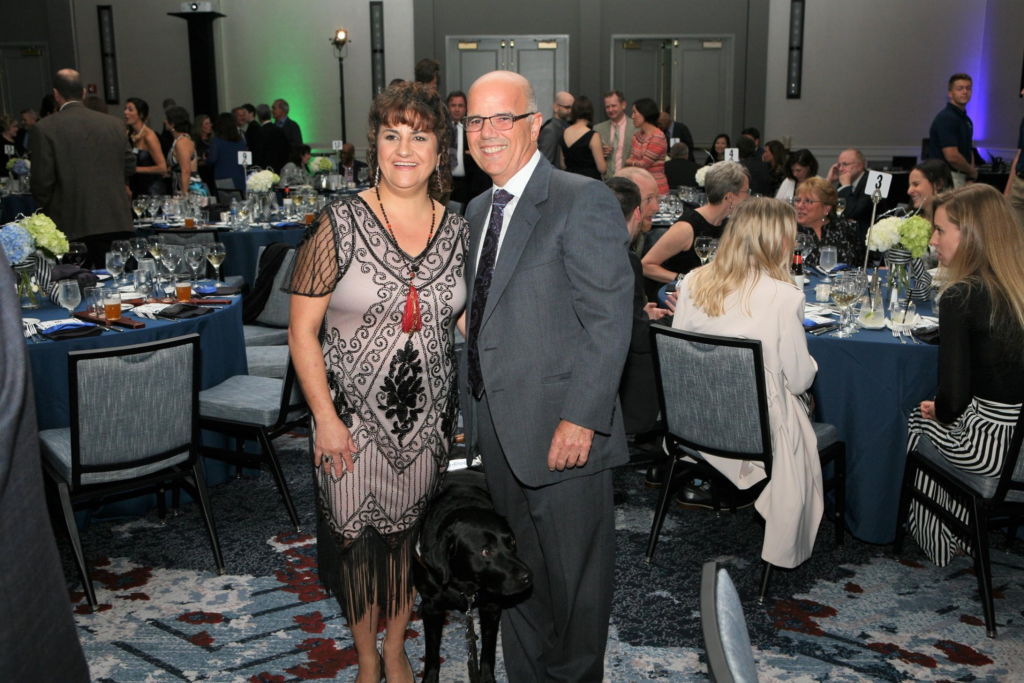
Orly and Amit nicely dressed at Dinner in the Dark 2019
Orly Shamir and her husband Amit (and guide dog Regan at the 2019 Dinner in the Dark to benefit Hope in Focus (formally Sofia Sees Hope).

was a fight. That’s why I want to give back and give light to the darkness.”

It began with chronic pain and prescriptions for Percocet and Oxycontin that offered relief and a false sense of well-being. Domestic abuse led to living in a shelter with her children. Orly finally realized she needed to get off the pills, but she could not.

“It was the beginning of torture for several years. That’s why we have an epidemic with opiates. It takes a lot of strength and support, and I tried three times over a four-year period. It takes everything out of you to get to the other end and never look back.”

Childhood memories of simmering aromas of lemons, parsley, cilantro, peppers, tomatoes and all the spices helped get her to the other side.

She tweaked her mother’s recipe during one of her mom’s visits from Montreal to Orly’s home in Toronto. When her don’t-you-dare-mess-with-ingredients mom left the kitchen, she took the opportunity, with guilt-laced excitement, to add a few more to the pot. 

“Voila, my specialty Moroccan Salmon, the champion of my life was born! At dinner my mom raved about her wonderful fish, and all I did was smile while my soul did a happy dance.”

Years later her addiction took its toll with memory loss and less ability to perform skills. She realized that improving her cognition could be accomplished several ways, including by eating healthy food, especially fish like salmon with lots of omega-3s.

It’s fatty, versatile and widely available, which is why Orly says her dish is champion food for anyone on any recovery journey, whether from illness, addiction or the blues.

“Without knowing it, this spiritually comforting food became physical healing food.”

Blind Ambition

Orly is a gifted chef, a title-holding athlete (Italian and Canadian dragon-boat racing!), a financial analyst fluent in French and a motivational speaker. She has two sons and a transgender daughter, all in their 20s living in Canada. Orly, her husband of 12 years, Amit, and her guide dog, a 6-year-old Black Lab named Regan, live in Hollywood, Fla.

She just graduated from a Florida culinary school with her sights set on developing a YouTube cooking series and a recipe app.

To help finance her project, she applied in January for the Holman Prize for Blind Ambition by making a 90-second video about her project of combining culinary and cooking expertise with her inspirational recovery story.

“Do you know blind people are 40 percent more likely to develop addictions?” she tells viewers in her video. “And did you know proper nutrition is key to recovery? … Although I’ve been legally blind my whole life, I lost my soul’s true vision through opioid addiction and poor health.”

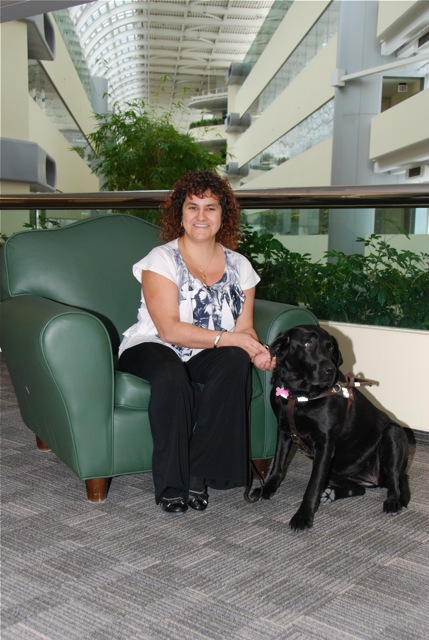
Orly sitting next to her black lab
Orly Shamir with her guide dog Regan.

Orly is turning her recovery story into a series of videos demonstrating healthy, delicious culinary delights, an accessible-to-all recipe app, and input from guest experts to help heal through the art of cooking.

She fashioned her simple and nutritious recovery recipes by using pronounceable ingredients, healthy fats, nuts and seeds and the like.

“It gives us more mental and physical strength because we start to feel better. We’re not as sluggish.”

Orly learned in March that she is one of 39 semi-finalists for the Holman Prize, selected from 109 applicants worldwide. Three winners will be selected in May.

The Holman Prize for Blind Ambition is annually awarded to three blind individuals to carry out a dream project to push limits and change perceptions about blindness around the world. The prize honors James Holman, a Victorian-era adventurer and author who became the first blind person to circumnavigate the globe; he also holds the distinction of being the most prolific traveler in history, sighted or not, prior to the invention of modern transportation.

Each Holman Prize winner receives up to $25,000 to fund a project or an adventure that will make an impact. The contest is sponsored by LightHouse for the Blind and Visually Impaired in San Francisco. The organization launched the prize concept to support the emerging adventurousness and can-do spirit of blind and low-vision people worldwide.

LightHouse CEO Bryan Bashin said the Holman Prize is not meant to save the world nor to congratulate someone for leaving the house. It is meant to change perceptions about what blind people can do.

“This prize will spark unanticipated accomplishments in the blindness community. You will see blind people doing things that surprise and perhaps even confuse you.” 

Previous winning projects include teaching blind people to become beekeepers in Uganda, hosting the first conference in Mexico for blind children and their families led by blind professionals, and recording a documentary series called “Planes, Trains & Canes” about navigating and accessing transportation systems in five cities around the world.

For the next step in the contest, Orly is creating an in-depth proposal due by the end of April. 

“My talent for cooking, my experiences all over the world tasting a plethora of inspiring favors, and my own story of failure, addiction and abuse woven in with courage, resilience and recovery – it was all for this.”

She Made Canadian Medical History When She Received a ‘Bionic Eye’

Orly Shamir made Canadian medical history when she became the first person in the country to receive a bionic eye. 

In a clinical trial six years ago, Dr. Robert Devenyi and his surgical team at a Toronto hospital implanted an Argus Retinal Prosthesis System (Argus II) onto the surface of Orly’s retina.

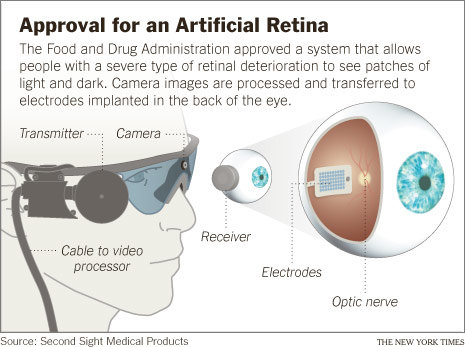
The Argus II, known as the bionic eye, stimulates the eye with electrodes to transmit visual information captured by a video camera to the patient’s brain. Toronto Western Hospital-University Health Network and Foundation Fighting Blindness provided support as collaborators on the 2014 clinical trial. California-based Second Sight developed the device.

Health Canada approved the system in 2015. It was approved in the United States in 2014 and in Europe in 2011. 

Orly has a form of Leber congenital amaurosis called LCA4 caused by a mutation in her AIPL1 gene. She could read large print as a child, but her vision deteriorated to minimal light perception and shadows in her 20s. Now 52, Orly took part in the 2014 trial when she was 46.

Photoreceptors in a healthy retina convert light into electrochemical pulses sent through the optic nerve and into the brain where they are decoded into images. When photoreceptors don’t function properly, as in the case of people with LCA, the first step in the vision process is disrupted and cannot transform light into images.

Approval for an Artificial Retina graphic

The Argus II bypasses damaged photoreceptors through electrodes implanted on the retina. Following the delicate three-hour surgery, Orly wore glasses containing a tiny camera that converted video images into a series of small electrical impulses transmitted wirelessly to the electrodes. Visual information transmits to the brain’s optic nerve when the pulses stimulate the retina’s remaining cells.

The visual improvement does not equal regular sight, but it allows patients to perceive light patterns, observe whether doors and windows are open, or pick up a glass.

Orly worked with rehabilitation experts, low-vision therapists and consultants from Second Sight. Three months following the surgery, she could detect contrast and recognize the difference between white and black. Because patients receive a form of artificial vision through this bionic eye, they need to re-train their brains through rehabilitation to learn and understand messages sent by the device. It’s kind of like learning a new language.

Orly volunteered to take part in the clinical trial for a year but presently does not use the technology. 

“I worked hard for two years, then decided to not use the system for personal use, as it didn’t provide any benefits as of yet.” 

Orly’s participation is exactly what a clinical trial is all about. She committed to making frequent visits and underwent testing over the course of a year. She knew that the goal of a clinical trial is to see whether a potential treatment is safe and effective, and that, while a trial participant might benefit, the trial’s purpose is to determine if the therapy works. 

“I was prepared in being part of advancing technology for blind/visually impaired people in the future. That’s always who I am,” she said. “I’m 100 percent ready and available to get updates and keep trying.”

All About Clinical Trials

Clinical trials are never done in a vacuum, or in a medieval basement where Dr. Frederick Frankenstein (pronounced Fronkensteen), his pretty lab assistant, Inga, and faithful houseboy, Igor, create a monster. 

While the creature from the 1974 movie “Young Frankenstein” turns out to be somewhat civilized, (see Gene Wilder as the young doctor and Peter Boyle as the monster doing the soft-shoe in white tie and black tails to “Puttin’ on the Ritz”), a caption above that lab team in a slide shown at the Hope in Focus (formally Sofia Sees Hope) second LCA Family Conference cautions: “Regulatory Oversight is Critical!”

Humor credit goes to Ben Shaberman, Senior Director of Scientific Outreach and Community Engagement for Foundation Fighting Blindness. He moderated a four-member panel discussion in a session called “All About Clinical Trials” at the July conference in Philadelphia. Shaberman reports on retinal research for the Foundation’s print and electronic publications. He also presents scientific advancements at local and national events and enjoys working with constituents to help them understand their retinal disease and current research that may benefit them.

Making sure trials are safe

Dr. Wiley Chambers II, MD headshot
Dr. Wiley Chambers II, MD

Panelist Dr. Wiley A. Chambers, the U.S. Food and Drug Administration’s Supervisory Medical Officer in the Office of New Drugs, said terrible outcomes can happen in trials not approved and regulated by the FDA. 

Dr. Chambers previously cited a disastrous case in which a 77-year-old woman traveled to an alleged clinic to have “stem cells” injected in her eyes in the hope of a cure or at least help for her macular degeneration. The procedure entailed separating “stem cells” from the woman’s belly fat and injecting them into her eyes to supposedly regenerate tissue. Not covered by insurance and not federally regulated, the procedure cost almost $9,000 and caused her vision to badly deteriorate after her retinas peeled away from her eyes.

The FDA assigns an Investigational New Drug (IND) number to every legitimate clinical trial.

“Just be careful when you go to a clinical trial or a physician that it is a legitimate trial,” Dr. Chambers said. “If they can’t give you an IND number, then walk away.” 

The website Clinicaltrials.gov includes trials that do have an IND number and unregulated trials that do not have an IND number. Avoid them. One must specifically ask if they have been issued an IND number, signaling the federal OK to proceed with clinical studies that happen in three phases.

Dr. Chambers also noted that trials are not for the benefit of participating patients. At the heart of a clinical trial is that researchers do not know what’s going to happen. A key word here is equipoise – a balance or counterbalance – of something. 

“We do the clinical trial and find out, does this have some efficacy and is it safe?”

Clinical trials tell as much information as possible, balanced with a doable study and doable patient commitment.

“It’s a balancing act,” he said. “The trials are to help inform people afterward.”

Research studies in humans are for potential therapies; they take 10 years or more to complete and can cost tens or hundreds of millions of dollars.

It took 12 years and $500 million to research and develop LUXTURNA™, an engineered virus delivered by subretinal injections of the human RPE65 gene, a gene that, when mutated, causes a form of LCA called LCA2 (RPE65-LCA). 

Tami in a pink shirt and Michael besides her in a bright orange shirt sitting at the 2019 LCA Family Conference
Tami and Michael Morehouse

Panel members Tami Morehouse and her husband, Michael, experienced firsthand what it feels like to take part in a Phase 1/2 LCA/RPE65 gene therapy trial. Intervention at earlier ages can offer better results with degenerative diseases like LCA and other inherited retinal diseases (IRDs) because photoreceptors diminish with age. 

The Cleveland-area couple talked about Tami’s pioneering experience as the oldest person, at 44, to take part in the LUXTURNA trial and regain some vision.

“It’s a huge, life-changing event for us,” Michael said.

Walking in Philadelphia three days after her first injection, Tami asked her husband if a building up ahead had stripes on it. Turns out, it was a parking garage with spaces between levels appearing as solid stripes. Before the injection, she could not see any part of the parking deck; her ability to discern solids and stripes meant the treatment already began improving her vision.

Tami had some vision in college and walked across the graduation stage by herself.  Michael described her progressive vision loss as, “Go on, go on, big drop; go on, go on, big drop; no diagnosis.

“She was on a path to darkness and she knew it,” he said.

He heard Jean Bennett, MD, PhD, on a Sunday radio show talking about her retinal research on the emerging therapy that came to fruition as LUXTURNA. He called her office early the next day, and said to the audience, “Guess who answered the phone?” (Here’s a link to a story on Dr. Bennett’s presentation at this conference.)  

Seven months later they received an email from Dr. Bennett, with the subject line: “Are you ready?”

Dr. Bennett conducted the studies with her partner in research and in marriage, Dr. Albert M. Maguire.  

In answer to a question about what she was thinking before beginning the trial, Tami said, “I don’t want to go totally blind.”

“In all honesty, I never thought that I’d ever have a shot at seeing … I kind of underestimated my possibilities. I became a much more functional person in day-to-day living. I would see who’s approaching. See my kids, my (softball-playing) daughter dancing off third base, taunting her catcher. 

“I got way more than I anticipated.”

Tami is part of the Sofia Sees Hope Family Connections program, calming fears and sharing her experiences with many patients along the way, including very young ones. 

“Even though huge strides have been made in clinical trials, they’re very frightened, they’re very nervous. 

“Jean and Albert explained so much; they relieved my fears in such a way that I’ve been able to transmit that to families, to moms. It’s a lot of pressure for some moms. 

“Just being able to say that it won’t hurt when Dr. Maguire inserts a needle in your eye because you’re out like a light; that is a beautiful thing because you don’t even know what’s happening to you.”

Tami advised potential surgical patients and their parents that even though undergoing surgery has the potential to do such good by improving vision, they should think about their expectations. “We are comfortable in our own zone; give us a little bit of change and it can throw us off.”

“If mom and dad or older siblings are calmer, the children are going to be more comfortable, too.”

Panelist Dr. Michel Michaelides, a founding member and head of clinical ophthalmology at MeiraGTx based in New York City and London, said clinical trials impact the daily lives of study participants because they’re required to undergo multiple tests, many of which, he said, are boring and tedious.

“We spend a lot of time letting people know what they’re really in for.”

Black and White headshot of Dr. Michel Michaelides
Dr. Michel Michaelides, a Founding Member and Head of Clinical Ophthalmology at MeiraGTx

Dr. Michaelides is the Principal Investigator of four interventional clinical trials and has 10 ongoing ethically approved studies. He is Professor of Ophthalmology at London’s UCL Institute of Ophthalmology in the Department of Genetics and Molecular Therapy, and Consultant Ophthalmologist at Moorfields Eye Hospital in the Departments of Inherited Eye Disease, Medical Retina and Pedriatric Ophthalmology.

Moderator Shaberman asked him what he says about therapy options to people who have lost a lot of vision. In advanced cases of vision loss, Dr. Michaelides said, optogenetics might be a relevant course of action. Optogenetics is the science of making cells in the retina that do not normally detect light, become light sensitive, and thereby aim to replace the lost light-sensitive cells (rods and cones/photoreceptors).

“The idea is to make cells that are not light sensitive, (be) light sensitive.”

Another course is retinal implant technology, also known as the “Bionic Eye” or “Artificial Vision,” in which doctors insert a light-sensitive microchip into the retina to provide a way to detect light. These implants can be placed on the surface of the retina (epi-retinal implants) or underneath the retina (sub-retinal implants). He also suggested the use of internal or external cameras with these implants, saying, “I think there’s going to be greater development in that area.”

Another avenue is stem cell therapies (cell therapies), in which donor cells could be used to grow fresh retinal cells for transplantation into the eye to replace lost cells. 

He also is involved in the development of therapies using stem cells to replace lost light-detecting retinal cells. He has been Principal Investigator of the first ocular stem cell therapy trial, which involved transplanting retinal pigment epithelial cells (non-light-detecting cells) in patients with advanced Stargardt Disease

His clinical trial for a potential LCA4 (AIPL1-LCA) therapy currently is recruiting participants.

Dr. Michaelides’ ocular research comprises 300 peer-reviewed publications and 25 book chapters. One of those research papers came into focus at the LCA conference in July, where an audience member – a mother of a child with LCA2 (RPE65-LCA) – told her story.

“Even after an ERG (electroretinography),*” she said, “nobody told us it was LCA.” 

She said reading one of his research publications on RPE65 put her family on the right track.

“That’s what we took to our doctor,” she said. “So, you’ve been really important to our journey.”